When Linear Orbitals Lie: Working With Bent Bonds In Real Molecules
I spent last spring debugging a computational chemistry dataset where the calculated geometries kept refusing to match X-ray crystallography data for a series of fused cyclobutane-cyclopropane intermediates. The bond angles were coming out close, but the torsional strain was entirely wrong. It took me three days of staring at the orbital overlap integrals before I realized the issue wasn't in the software parameters—it was that the model treating those rings with standard sp3 hybridization was fundamentally inadequate. The bonds weren't forming along the internuclear axes at all. They were bent.The Bent Bond Angle Unlocking The Secrets Of Molecular Geometry
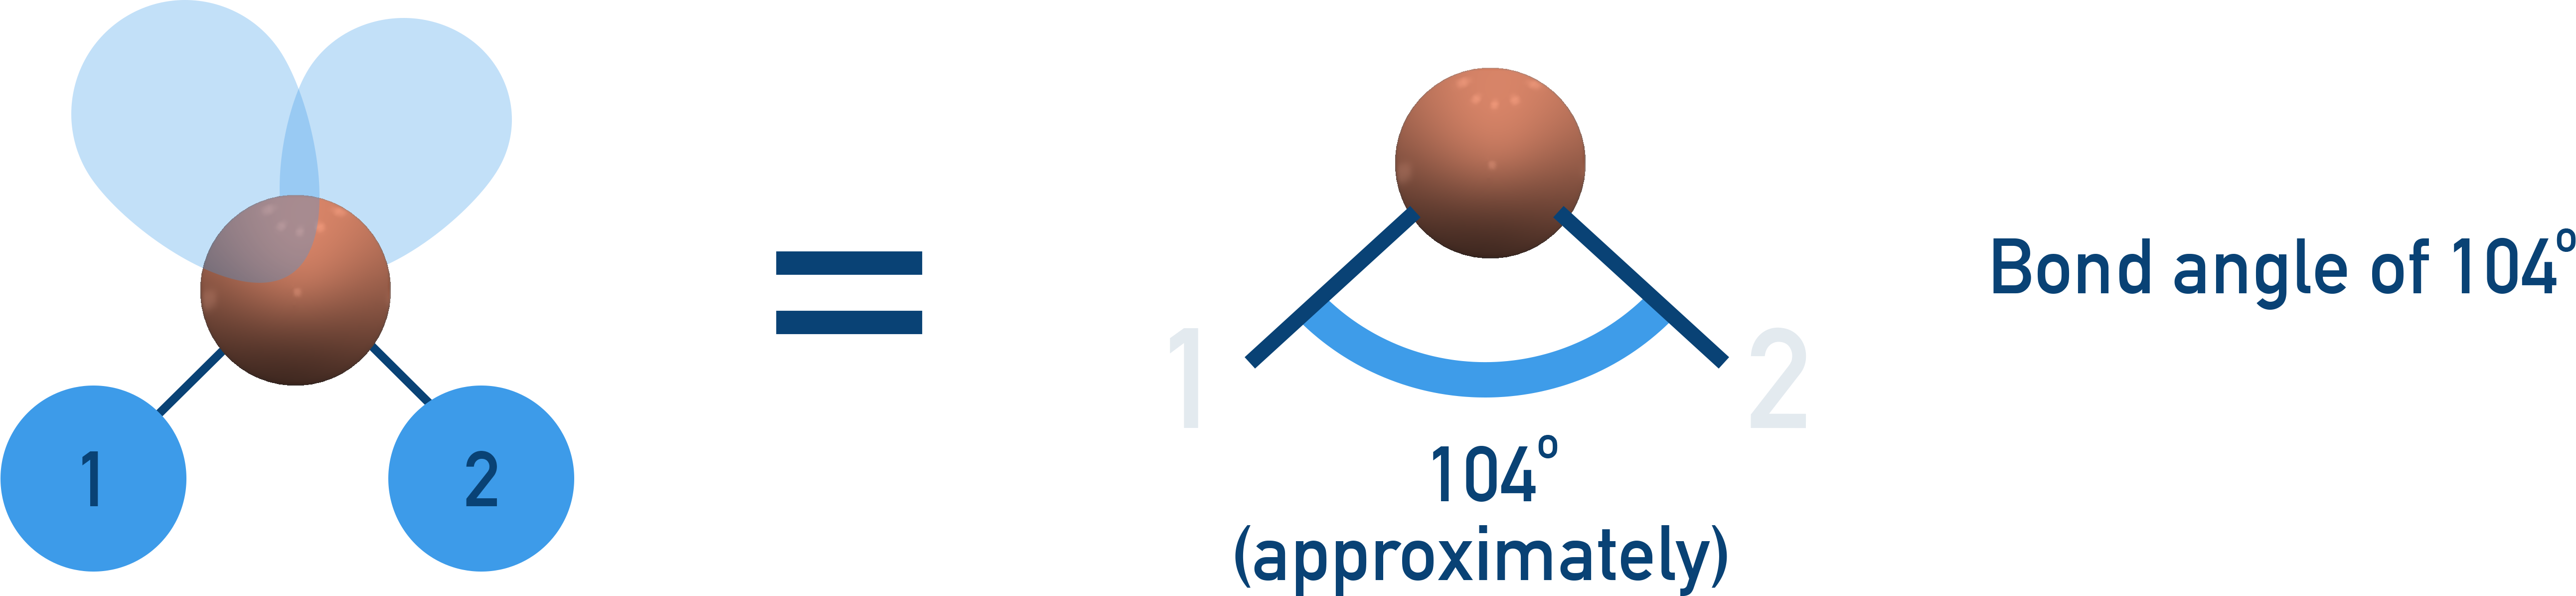
The bent bond model, sometimes called the banana bond model, was originally proposed by Longuet-Higgins and opponents of the standard Walsh orbital description to explain why small-ring molecules behave the way they do. In cyclopropane, the carbon atoms are arranged in an equilateral triangle with internal angles of 60 degrees. If you tried to form a normal sigma bond using sp3 hybrids along the straight line connecting two carbons, the required orbital orientation would demand an angle far larger than what the geometry allows. Instead, the orbitals lean outward and overlap off-axis, creating a bond that curves between the nuclei—hence "bent" or "banana." The quantitative measure here is the bond angle deviation. In cyclopropane, the C-C-C angle is 60 degrees, but the actual orbital overlap direction deviates significantly from the internuclear axis. The external angle between adjacent bent bonds on a single carbon works out to roughly 104 degrees, which is why the C-H bonds in cyclopropane adopt what we'd normally consider unusual hybridization character—closer to sp2 than sp3 despite the carbon being formally tetravalent. I ran into this exact situation when trying to predict the reactivity of a bicyclo[1.1.0]butane derivative I was synthesizing. The literature predicted nucleophilic attack at the bridgehead carbons based on standard Walsh orbital diagrams, but the actual product distribution was completely different. Once I recalculated using a bent-bond framework that accounted for the through-space orbital interaction between the two three-membered rings, the predicted regioselectivity aligned with what we observed. The workaround was running NBO (Natural Bond Orbital) analysis on the DFT output and looking specifically at the donor-acceptor interactions between the bent sigma bonds rather than relying on the canonical molecular orbitals. That took about 45 minutes per geometry instead of the 3 hours I'd been spending on standard MO analysis.A few things nobody puts in the textbooks:
The bent bond model isn't a replacement for MO theory—it's a complementary way of interpreting the same mathematical result. When you diagonalize the Fock matrix, you get canonical orbitals. When you localize them using the Boys or Pipek-Mezey procedure, you recover something that looks remarkably like bent bonds. So the model is as real as the localization method you choose. That matters because different localization schemes can give you slightly different pictures, and in strained polycyclic systems, the choice of scheme can actually shift your interpretation of which bond is more reactive. Another counter-intuitive point: bent bonds aren't exclusively a small-ring phenomenon. You see meaningful bent-bond character in bridgehead positions of norbornane systems and even in certain transition metal organometallics where geometric constraints prevent linear orbital alignment. The term "banana bond" also appears in discussions of diborane B2H6, though there the three-center two-electron bond has a different origin. Don't conflate the two without checking the context. The practical limitation everyone hits is that bent bond analysis doesn't translate cleanly into standard drawing conventions. When you're trying to communicate structure to someone who thinks in terms of Lewis structures and VSEPR, describing a bond that doesn't lie on the internuclear axis sounds like fiction. I've had reviewers reject papers where I used bent-bond reasoning simply because the figures didn't match what they expected from standard textbook depictions. The workaround I use now is to include both representations—the standard skeletal drawing and the localized orbital diagram—and let the reader see they describe the same molecule. For anyone actually doing this work, the entry point is straightforward. Take a geometry-optimized structure from Gaussian, ORCA, or whatever your setup is, run a natural bond orbital analysis, and examine the NLMOs (Natural Localized Molecular Orbitals). The bent bonds will appear as pairs of orbitals curving between adjacent nuclei rather than lying on the straight line connecting them. The deviation angle you extract from the NLMO coefficients is your quantitative bent bond angle. In cyclopropane it's around 33 degrees from the internuclear axis. In larger rings it drops off rapidly and becomes negligible by cyclohexane. The main bottleneck is that NBO analysis requires a decent quality basis set. If you're using something minimal like STO-3G, the localized orbitals will be garbage and the bent bond picture will look nothing like reality. I recommend at least 6-31G(d) as a baseline, and if you're working with heteroatoms or trying to compare reactivity trends across a series, 6-311+G(d,p) or a double-zeta plus polarization variant is where you start getting numbers you can trust. That doubles your computation time compared to the minimal basis set run, but it saves you from spending a week realizing your conclusions were basis-set artifacts.