Understanding Phase Separation in Liquid-Liquid Extraction
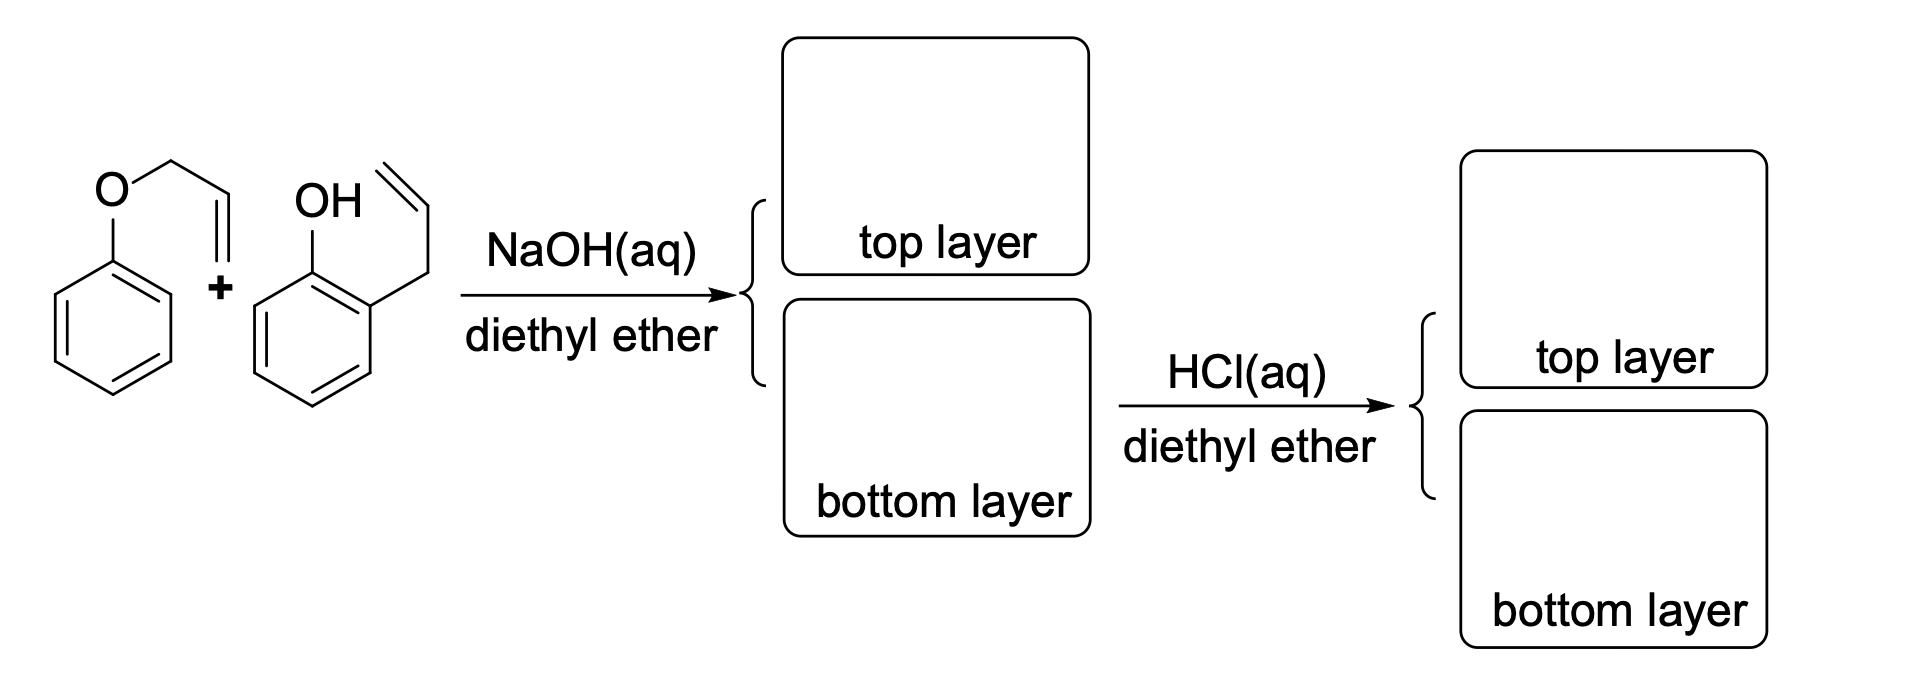
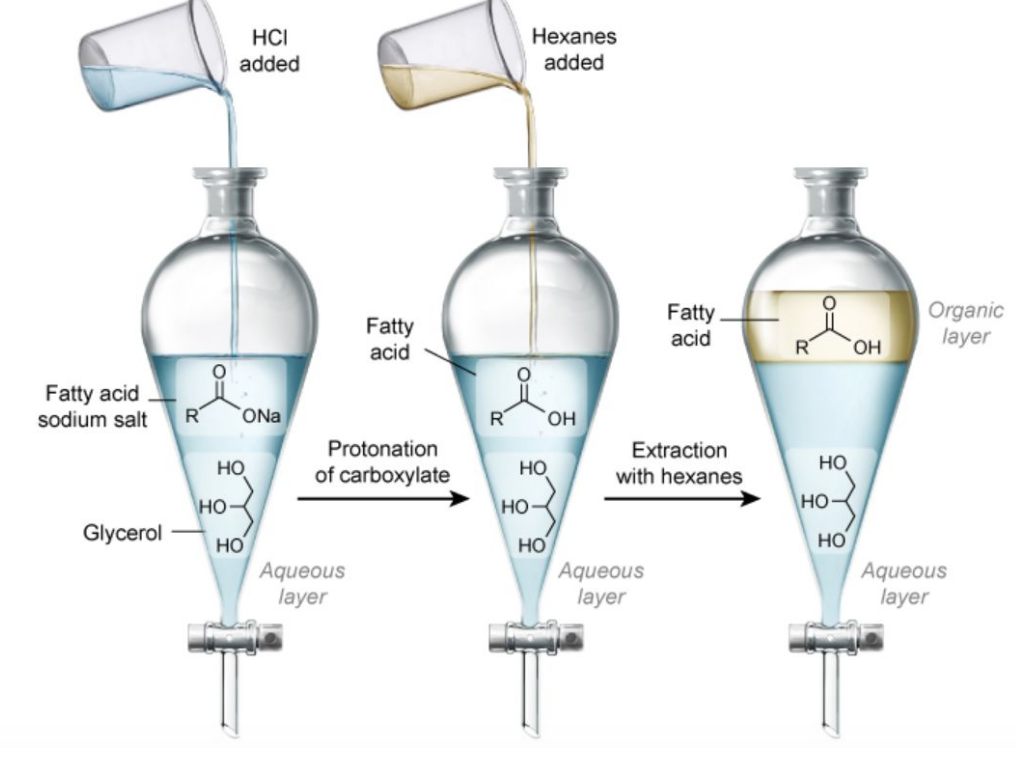
When you shake a separatory funnel containing water and an ether solvent, two distinct layers form. Getting them to separate cleanly and identifying which is which is one of those things that seems obvious in theory but causes problems in practice every single day. The aqueous layer is water-based. It contains dissolved salts, polar compounds, and anything hydrophilic. The ether layer (usually diethyl ether or MTBE in lab settings) is the organic phase. It pulls nonpolar and moderately polar organics out of the water. Simple enough on paper. The problem is identifying which layer is which when they look identical. Both are clear, colorless liquids. You add your extract, shake, let it sit, and suddenly you have two nearly indistinguishable phases sitting in a funnel.
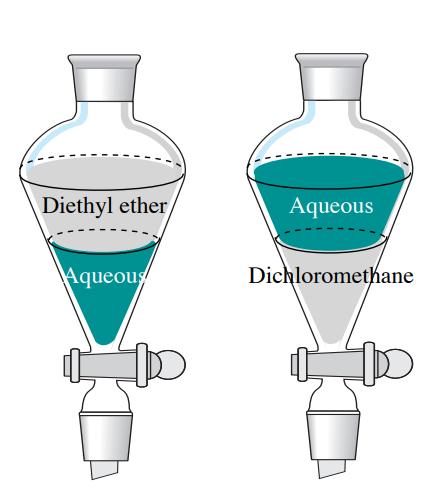
Here is the practical way to tell them apart: Add a few drops of water to the top of the mixture. Watch where the drops go. If they dissolve into the top layer, that top layer is aqueous. If the drops fall through the top layer and disperse into the bottom layer, the bottom layer is aqueous. This takes about ten seconds and saves you from accidentally discarding the wrong phase. I learned this the hard way during a graduate school run where I was extracting a modestly polar product from a reaction mixture. I assumed the bottom layer was aqueous because I had used a lot of brine wash, and I pipetted off what I thought was the organic layer. Turns out I had used dichloromethane as my solvent, which is denser than water, so the organic layer was on the bottom. I lost roughly four hours of product because I had not verified the layer identity before separating. That mistake has haunted me ever since.
Density is the deciding factor
Most ether solvents are less dense than water. Diethyl ether has a density around 0.71 g/mL. MTBE is about 0.74 g/mL. This means the organic layer typically sits on top of the aqueous layer. But there are important exceptions that trip people up regularly. Chloroform weighs 1.49 g/mL. Dichloromethane is 1.33 g/mL. If you are using either of these instead of ether, the organic layer is on the bottom. This is the most common reason people lose product during extractions. You see two layers, you assume the top is organic because that is what your textbook showed you, and you drain the wrong one out. Adding saturated sodium chloride (brine) increases the density of the aqueous layer to roughly 1.2 g/mL. This actually widens the density gap and makes separation cleaner, but it also means even solvents that are slightly heavier than pure water can still float on top of brine. Always check, not assume.
Get the Full Details

Emulsions are the real enemy
Sometimes the layers refuse to separate. An emulsion forms and you are left with a cloudy interface that looks like mayonnaise. This happens more often than most protocols acknowledge. Surfactants, fine particulates, proteins, and certain byproducts of organic reactions all stabilize emulsions. My standard approach when an emulsion appears: first try gentle swirling instead of shaking. Next, add a small amount of brine and mix again — the high ionic strength collapses many emulsions. If that fails, a pinch of sodium sulfate or a few drops of isopropanol usually breaks it. I have also used centrifugation at 3000 RPM for about three minutes when nothing else worked, though that requires transferring the entire mixture to centrifuge tubes, which is another opportunity to lose material. The one case where I consistently struggle is when working with plant extracts rich in lipids and waxes. Those materials form stubborn emulsions that can hold for hours. In those situations I skip the separatory funnel entirely and use a solvent system where the phases are more immiscible, or I filter through celite before attempting any separation.
Why incomplete separation ruins yields
When the interface between layers is dirty or cloudy, some of your product is trapped in that middle zone. Draining too aggressively pulls emulsion into your collected layer. Stopping too early leaves product behind in the funnel. The difference between a 78 percent recovery and a 92 percent recovery in a routine extraction often comes down to how carefully someone handles the interface. A practical technique: drain the bottom layer slowly and stop when the meniscus reaches the stopcock. Transfer the remaining top layer through the top of the funnel rather than draining it, which avoids pulling any residual interface material into your collection vessel. It adds maybe thirty seconds per extraction but prevents cross-contamination between phases.
When ether is the wrong choice
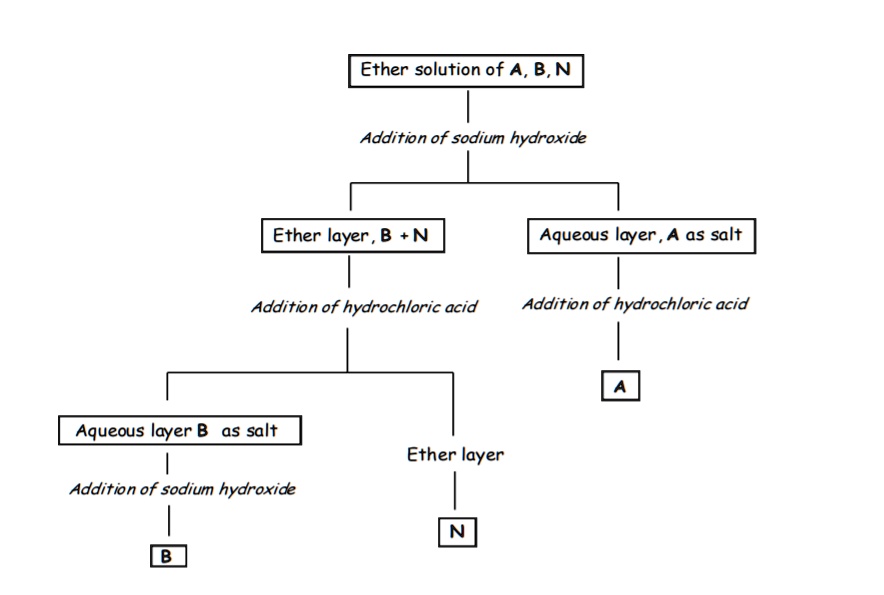
Diethyl ether has real drawbacks. It forms peroxides on storage, which becomes a safety issue if you are running multiple extractions over weeks. It is highly volatile, creating pressure buildup in a closed separatory funnel if you are not venting frequently. It has limited solvating power for anything more than moderately nonpolar compounds. For compounds that are quite nonpolar, hexanes or toluene work better. For moderately polar compounds that ether misses, ethyl acetate or dichloromethane are more appropriate. The solvent choice should match your target compound's polarity, not just tradition. Using ether for everything because it is convenient is how people end up with poor partition coefficients and frustrated colleagues. I also want to note that this method has clear limitations. It assumes your compound has significantly different solubility between the two phases. If the distribution coefficient is close to one, no amount of careful layer identification will give you good recovery. In those cases you need multiple extractions with fresh solvent, or you need to change the pH to ionize or neutralize the compound and shift its solubility. Extraction is not a magic bullet. It is a partitioning tool, and like any tool, it only works within its operating range.