What Actually Happens When You Mix Two Solutions
You pipette solution A into solution B, wait ten seconds, and get nothing. No precipitate, no color change, no gas bubbles. You check your notes and swear you balanced the equation correctly. This happens more often than you'd think, and it almost always comes down to not understanding what stays dissolved and what falls out. Solubility Guidelines The Key To Unlocking Chemical Reactions isn't about memorizing a chart and hoping for the best. It's about predicting whether your reaction will actually produce what you expect before you waste reagents and lab time on it. The rules themselves are straightforward, but applying them correctly requires knowing where the exceptions live.
Where to Start: Cations and Anions
Every solubility rule ties back to ion pairing. When you combine an aqueous solution containing a specific cation with another containing a specific anion, you're really asking whether that new ionic compound exceeds its Ksp in the given conditions. The standard solubility tables compress this into categories like "always soluble" or "usually insoluble," but those words hide important qualifiers. Here's how I approached this when I was still teaching undergrad labs. I stopped having students memorize the full table and instead made them predict three reactions each before running them. Sixty percent of the class got at least one wrong because they applied the nitrate rule blindly, forgetting that the rule only covers nitrate salts, not every compound containing nitrogen. A student once mixed ammonium sulfide with lead nitrate expecting a sulfide precipitate and was genuinely confused when the Ksp of lead sulfide meant the reaction went to completion almost instantly. That student learned the difference between qualitative prediction and quantitative reality faster than any lecture could teach them.
The Rules That Actually Matter
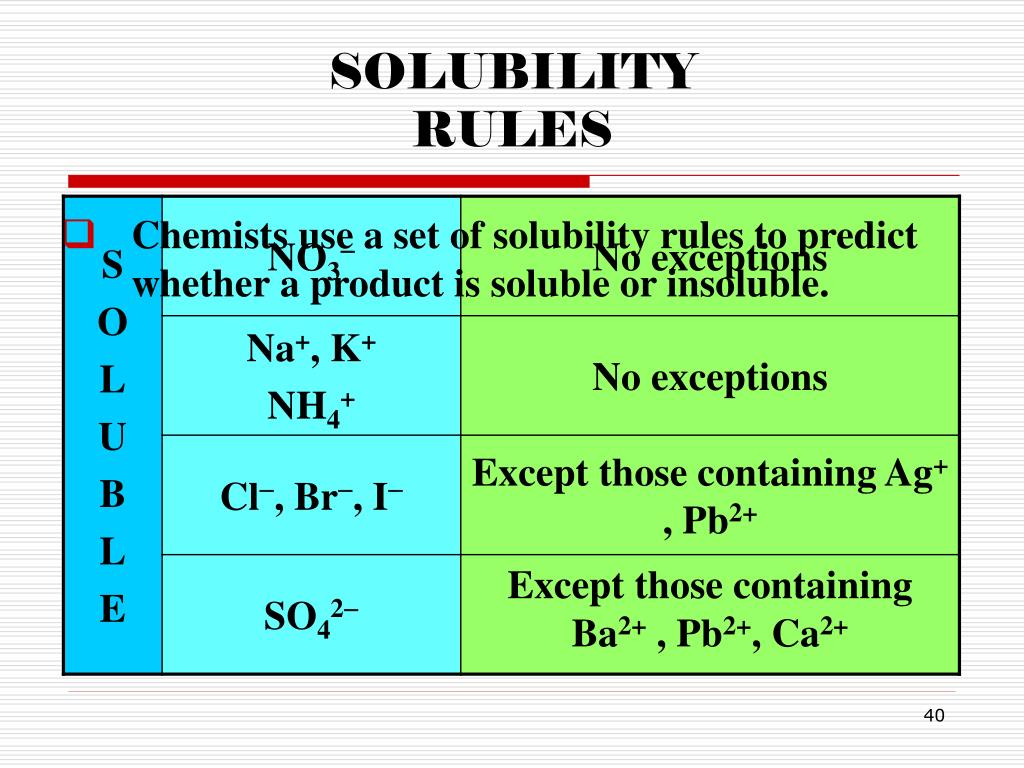
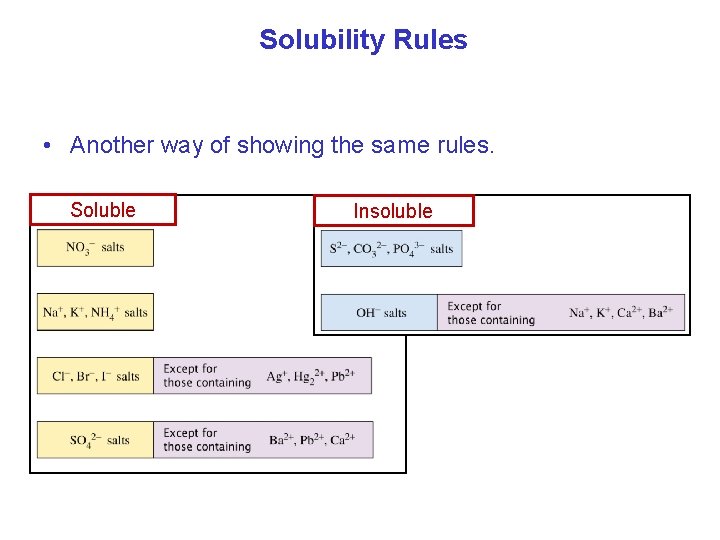
Soluble without exception: Group 1 cations (lithium, sodium, potassium, rubidium, cesium) and ammonium. Nitrates, acetates, and perchlorates. These dissolve regardless of what they're paired with under normal lab conditions. Mostly soluble: Chlorides, bromides, and iodides. The common exceptions are silver, lead(II), and mercury(I). If you're working with chloride salts and need to know whether precipitation will occur, check whether any of these three metals are in play. Fluoride follows a different pattern entirely — it precipitates with calcium, magnesium, and the heavier group 2 metals, which contradicts the general halide trend and catches people off guard regularly. Mostly insoluble: Hydroxides, carbonates, phosphates, sulfides, and chromates. The Group 1 and ammonium exceptions apply here too. Calcium hydroxide sits in a gray zone — it's technically sparingly soluble at about 1.73 g/L at 25°C, which means saturated solutions form clouds of precipitate that look like complete failure but are actually just at equilibrium. Students routinely discard what they think is a failed synthesis when they've simply created a saturated slurry.
Get the Full Details
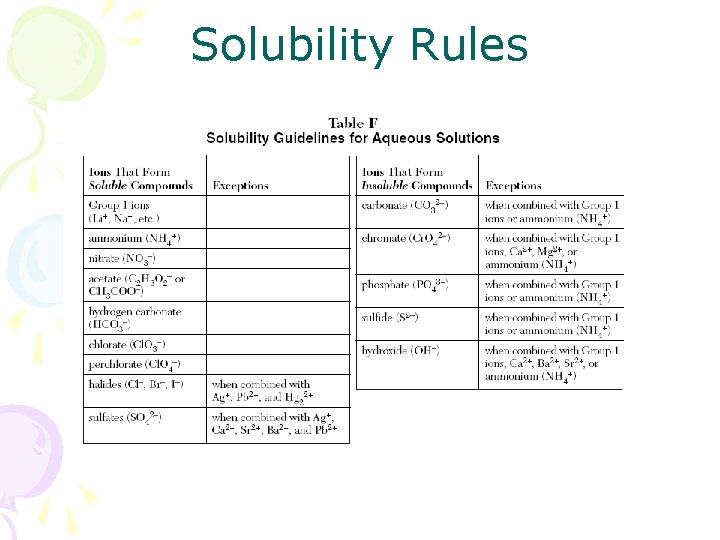
Sulfates: Generally soluble. The exceptions are calcium, strontium, barium, lead, and mercury(II). Barium sulfate is essentially insoluble — Ksp around 1.1 × 10¹ — which is why it's used in medical imaging. If you're doing qualitative analysis and add sulfate to an unknown, a white precipitate strongly suggests barium, strontium, or lead. Not definitive on its own, but it narrows the field quickly.
Temperature Changes Everything
The solubility tables you're looking at were measured at 25°C. Most textbooks don't emphasize this enough. Potassium nitrate, for example, has a solubility of about 32 g per 100 mL of water at 25°C, but that jumps to roughly 247 g per 100 mL at 100°C. If you're trying to crystallize KNO by cooling a hot saturated solution, you recover most of it. If you try the same thing with NaCl, you recover almost nothing because its solubility barely changes with temperature — about 36 g/100 mL at 25°C and 39 g/100 mL at 100°C. This distinction determines whether recrystallization is even a viable purification method. I once had a graduate student spend three days trying to purify a compound by recrystallization from water, only to discover her product had near-zero temperature-dependent solubility. She should have tested solubility at two temperatures before committing to the procedure. A five-minute test — dissolve a pinch in cold water, heat it, observe — would have saved her the entire effort.
Common Pitfalls That Waste Lab Time
Pitfall one: assuming "insoluble" means completely insoluble. Every precipitate has a Ksp. Silver chloride's Ksp is 1.8 × 10¹, which translates to about 1.3 mg per liter of water still dissolved after precipitation. In a typical undergraduate experiment where you're mixing 0.1 M solutions, the amount remaining in solution is negligible for most purposes. But if you're doing analytical work or trace metal analysis, that residual concentration matters. I've seen students report near-100% recovery from a supposedly quantitative precipitation and then wonder why their gravimetric results were consistently low. The missing mass wasn't lost — it was still dissolved. Pitfall two: ignoring complex ion formation. This is the one that quietly ruins otherwise correct predictions. Add ammonia to silver chloride precipitate and it dissolves. Not because the solubility rules changed, but because Ag forms the diamminesilver(I) complex, [Ag(NH)], with a formation constant around 1.7 × 10. The effective solubility of AgCl in 1 M ammonia is roughly a thousand times higher than in pure water. Similarly, zinc hydroxide redissolves in excess NaOH to form [Zn(OH)]². If you're doing qualitative analysis and your precipitate disappears when you add what should be a further reactant, check for complexation before declaring your procedure flawed. Pitfall three: pH dependence of anions. Carbonate, phosphate, sulfide, and hydroxide solubilities are all pH-dependent. Add acid to a carbonate precipitate and it dissolves because CO² gets protonated to HCO and then HCO, pulling the dissolution equilibrium forward. Sulfide precipitates behave similarly — HS is a weak acid, so lowering pH increases sulfide solubility significantly. In qualitative analysis schemes, this is deliberately exploited to separate groups of cations. Group 2 sulfides precipitate in acidic solution while Group 4 sulfides require basic conditions. If you skip the pH control, your separation fails and you get crossover contamination.
A Specific Problem I Encountered
I was running a synthesis where I needed to precipitate calcium oxalate from a solution that also contained significant magnesium. The solubility rules say both calcium oxalate and magnesium oxalate are insoluble, but their Ksp values are close enough that selective precipitation is theoretically possible. Calcium oxalate Ksp is roughly 2.7 × 10 while magnesium oxalate is about 8.6 × 10. The difference is about four orders of magnitude, which should be plenty. What I hadn't accounted for was the effect of ionic strength on activity coefficients. At the concentrations I was working with, the mean activity coefficient dropped enough that the effective solubility product shifted. Magnesium oxalate co-precipitated at levels high enough to contaminate the product. The workaround was to add the oxalate slowly with vigorous stirring while maintaining a slightly acidic pH (around 4) to suppress magnesium oxalate formation, then adjust to neutral after the calcium had fully precipitated. I confirmed purity by testing the filtrate for magnesium with ammonium phosphate — no precipitate meant the magnesium stayed in solution as intended. This added about twenty minutes to the procedure but eliminated a purification step that would have taken hours.
How to Actually Use These Rules in Practice
Don't just read the table. Write out the full ionic equation before you mix anything. Break every soluble compound into its constituent ions. Then recombine the ions in every possible pairing and check solubility for each combination. This catches double displacement reactions where the driving force isn't the one you initially assumed. I've seen people predict no reaction because they focused on the obvious salt product while missing that a weak electrolyte or gas was also forming. When in doubt, look up the Ksp rather than relying on the qualitative label. "Sparingly soluble" covers a range from Ksp = 10³ to 10¹² depending on the source. If you need to know whether precipitation will actually occur at your concentrations, calculate the ion product Q and compare it to Ksp. If Q exceeds Ksp, precipitation happens. If Q is below Ksp, nothing visible occurs regardless of what the general rule says. This calculation takes thirty seconds and prevents hours of confused troubleshooting later. For quick reference: Keep a Ksp table open while you work. The CRC Handbook of Chemistry and Physics has one that covers the most common compounds. Free online databases like the NIST Chemistry WebBook also list solubility data with temperature dependence, which is useful when your reaction doesn't run at 25°C. I print a single-page summary and tape it to my fume hood. It saves me from opening three different references when I'm mid-experiment.
When Solubility Rules Fail Completely
They don't account for organic solvent mixtures. Add ethanol to an aqueous ionic solution and solubility drops dramatically for most salts. This is the basis for many precipitation-based isolation methods, but the rules you learned in gen chem don't cover it. If you're working with partial aqueous-organic systems, you need experimental solubility data, not textbook heuristics. They also ignore kinetic factors. Some precipitates form slowly or create supersaturated solutions that resist nucleation for hours or days. Sodium acetate is the classic example — it's extremely soluble and can form stable supersaturated solutions that precipitate explosively when seeded. The solubility rules would tell you nothing about this behavior. If your reaction seems like it should precipitate but doesn't, give it time, scratch the glassware with a stirring rod, or add a seed crystal before concluding the prediction was wrong. Understanding solubility guidelines isn't about having perfect predictions. It's about having a starting point that lets you anticipate what might happen, recognize when something unexpected is occurring, and know which variable to adjust when your reaction doesn't behave as the simple rules suggest.