Figuring Out Which Pathway Your Reaction Takes
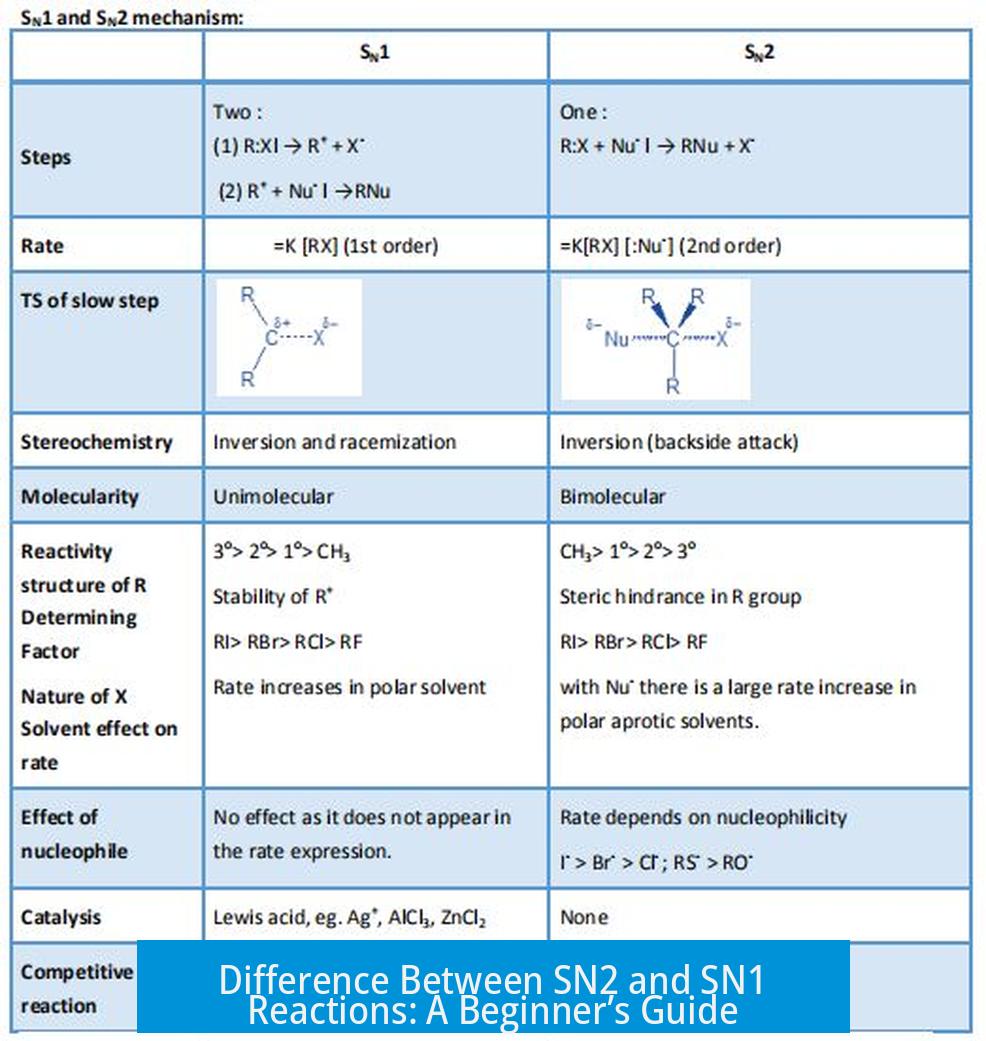
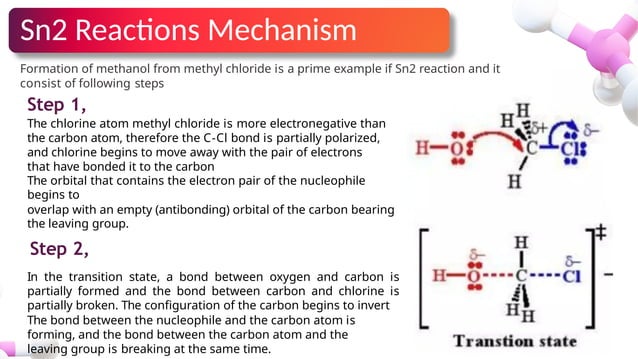
Most students learn SN2 as the backside attack and SN1 as the carbocation route, then spend hours memorizing flowcharts that fall apart the moment they look at a real lab notebook. The truth is less clean, and honestly, it is more useful to understand the signals you actually see. I spent three years running these reactions before the patterns stopped feeling like guesswork. Here is how I approach it now. The method starts with the substrate, not the nucleophile. A primary carbon with a good leaving group almost always goes SN2 unless something pushes it otherwise. Tertiary substrates are essentially locked into SN1 or E1 territory. Secondary carbons are where everything gets interesting and where people waste the most time. I used to treat every secondary halide as a coin flip until I learned to read the solvent and temperature clues. Solvent choice is the signal most people ignore. Polar aprotic solvents like DMSO, DMF, and acetonitrile accelerate SN2 dramatically because they leave the nucleophile relatively unsolvated. Polar protic solvents like water, methanol, and ethanol stabilize carbocations through hydrogen bonding and therefore favor SN1. This is textbook, but the practical detail is the magnitude. Moving from ethanol to DMSO for a secondary substrate can shift the product ratio from a messy 60-40 mixture to something cleanly SN2-dominated in under two hours at room temperature.
Nucleophile strength matters too, but not in the way beginners expect. A strong nucleophile does not guarantee SN2 if the substrate is tertiary. I once ran a reaction trying to force an SN2 on a neopentyl-like system using a very strong alkoxide in DMSO. The reaction did not proceed through substitution at all. Elimination dominated, and I got mostly alkene product after four hours. Switching to a weaker, bulkier nucleophile and lowering the temperature to 0 degrees Celsius changed nothing about the outcome. The substrate geometry itself was the bottleneck. I ended up using a different synthetic route altogether, going through an mesylate with a thiol nucleophile under milder conditions, which gave me the substitution product in about 78 percent yield over six hours. Temperature is another practical lever. SN1 reactions have higher activation energies for the rate-determining ionization step, so warming the reaction generally favors unimolecular pathways. Cooling tends to suppress carbocation formation and can tilt secondary systems toward bimolecular displacement. I usually run suspicious secondary substrates at room temperature first, then if the NMR shows elimination or racemization products, I drop it to reflux for SN1 or cool to 0°C for SN2, depending on what I want. Leaving group ability is non-negotiable for both pathways, but the threshold is lower for SN2. Iodide, tosylate, and mesylate work reliably. Bromide is acceptable but slower. Chloride is sluggish without activation, and fluoride is essentially inert for substitution under standard conditions. For SN1, even chloride can work if the resulting carbocation is stable enough, which is why tertiary chlorides sometimes ionize in wet acetone while primary chlorides sit there doing nothing.
Concentration is the detail that separates people who predict mechanisms from people who actually control them. SN2 is second-order, meaning the rate depends on both substrate and nucleophile concentration. Doubling the nucleophile concentration doubles the rate. SN1 is first-order in substrate only. If you are trying to push a borderline secondary system toward SN2, running it at high nucleophile concentration, ideally 2 to 3 molar, is more effective than almost anything else I have tried. It is a simple trick that most lab manuals do not emphasize enough. There is a misconception about stereospecificity that causes real problems. SN2 inverts stereochemistry, and SN1 racemizes it. That is true in ideal cases. In practice, I have seen SN1 reactions produce mostly inverted products when the leaving group blocks one face of the carbocation, creating what is called ion pairing. The anion stays close to one side of the intermediate, so the nucleophile attacks from the opposite side more often than a truly random model predicts. I encountered this with a secondary brosylate in aqueous acetone. The product showed 70 percent inversion instead of the 50-50 racemic mixture the textbooks imply. Running the same reaction with a non-nucleophilic counterion changed the ratio closer to racemic, confirming the ion-pairing effect. Another nuance beginners miss is that SN1 and E1 are not separate competing pathways in the way people treat them. They share the same rate-determining step: carbocation formation. Once the carbocation exists, it can lose a proton to give an alkene or capture a nucleophile to give substitution. The ratio between them depends on the nucleophile's basicity and the temperature. Stronger bases favor elimination. Higher temperatures favor elimination because it has a higher activation energy. I typically see E1 products dominate above 80°C in solvolysis reactions, even when I am trying to get substitution.
Get the Full Details
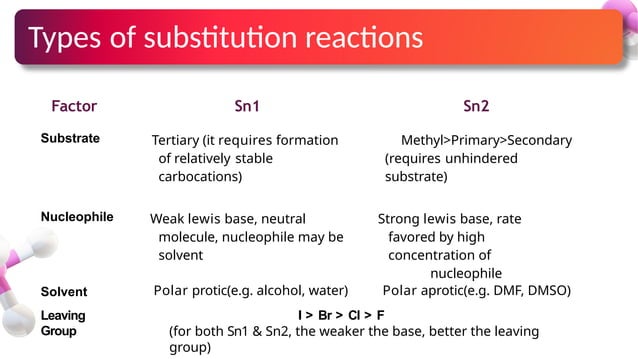
SN2 and E2 competition follows a similar logic. Strong, bulky bases like potassium tert-butoxide favor elimination over substitution even on primary substrates. Strong, small bases like sodium methoxide or sodium ethoxide still give mostly SN2 on primary carbons but start shifting toward E2 on secondary carbons. I use this to my advantage when I need an alkene, but it costs me time purifying elimination byproducts when substitution was the goal. Here is a practical decision sequence I follow before setting up any substitution reaction. Identify the substrate class. Primary means SN2 unless the beta carbon is highly branched. Tertiary means SN1 or E1, possibly E2 with strong bases. Secondary means look at the solvent, nucleophile, temperature, and concentration. If the solvent is polar aprotic and the nucleophile is strong and small, SN2 is likely. If the solvent is polar protic and the nucleophile is weak or the solvent is also the nucleophile, SN1 is more likely. Check the leaving group. If it is poor, consider converting it first. Run a small test at room temperature and analyze by TLC or GC before committing to a full scale. One thing this framework does not handle well is allylic and benzylic substrates. They can undergo SN1 through stabilized carbocations even when they are primary. They also undergo SN2 faster than ordinary primary substrates because the transition state is stabilized by conjugation. I learned this the hard way when a primary allylic chloride gave a racemic substitution product in ethanol at room temperature. The carbocation intermediate was clearly formed despite the primary classification. The workaround was using a non-ionizing solvent like acetonitrile with a strong nucleophile, which forced the SN2 pathway and gave clean inversion in 85 percent yield.
The biggest limitation of this approach is that it predicts tendencies, not certainties. Borderline secondary systems in mixed solvent conditions can give product distributions that vary significantly between batches due to moisture content, trace impurities, and subtle temperature gradients in the reaction vessel. I have seen the same reaction run on two consecutive days give 60 percent SN2 on day one and 40 percent SN2 on day two because the lab humidity changed and the solvent absorbed enough water to tip the balance toward ionization. There is no workaround for this except rigorous solvent drying and consistent experimental conditions. If you are working with particularly stubborn substrates where neither pathway gives clean results, switching to a different leaving group or using phase-transfer catalysis with a crown ether can sometimes force the reaction down a more predictable route. I had a case where a secondary Tosylate in DMSO gave a 50-50 mixture of substitution and elimination products regardless of temperature. Adding 18-crown-6 and using potassium acetate as the nucleophile pushed the substitution yield to about 82 percent within three hours at room temperature. The crown ether sequestered the potassium cation and increased the effective nucleophilicity of the acetate without increasing basicity significantly. Understanding SN2 and SN1 is less about memorizing rules and more about learning to read the combination of substrate, nucleophile, solvent, temperature, and concentration as a single system. The individual factors matter less than how they interact. A good nucleophile in the wrong solvent will underperform. A perfect substrate in the wrong temperature range will give elimination instead of substitution. The reactions are predictable once you stop treating each variable in isolation and start looking at the whole picture.