What Actually Determines SN1 or SN2 When You're Standing at the Fume Hood
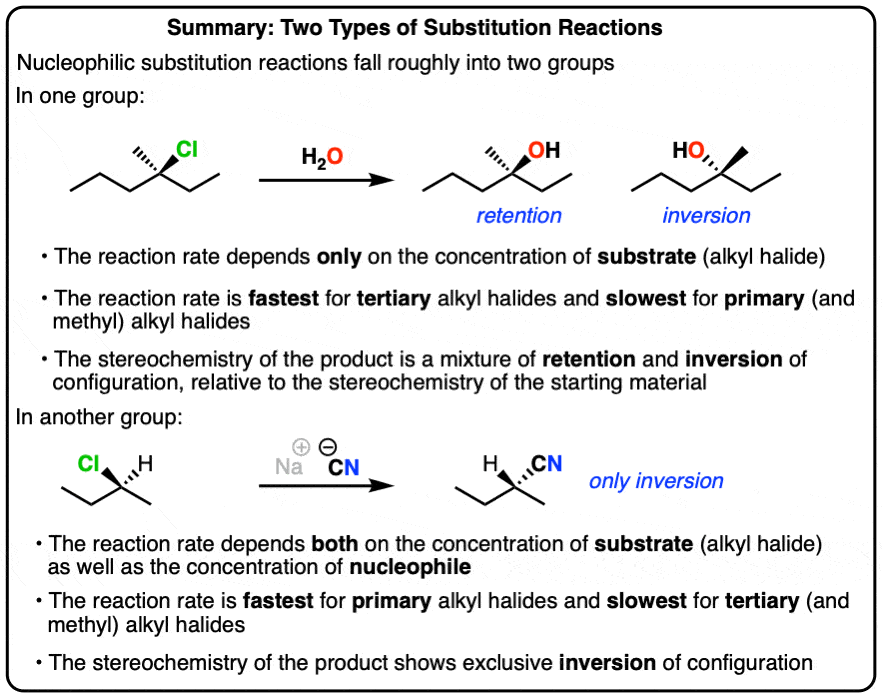
Most organic chemistry courses present SN1 and SN2 as clean, separate categories with predictable outcomes. That is not how it works in practice. I spent three semesters running substitution reactions and still get surprised by which pathway dominates. The real question is not whether a reaction is SN1 or SN2, it is what combination of substrate, nucleophile, solvent, and temperature will push it one way or the other, and whether you will get the product you actually want. Start with the substrate. This is the factor that matters most, and students routinely underestimate it. Primary carbons almost exclusively go SN2. Tertiary carbons almost exclusively go SN1. Secondary carbons are the problem zone. That is where you need to pay attention to everything else, because small changes flip the mechanism completely. The SN2 mechanism is one step. The nucleophile attacks the backside of the carbon at the same time the leaving group departs. There is no intermediate. The rate depends on both the substrate concentration and the nucleophile concentration, expressed as rate = k[substrate][nucleophile]. Inversion of configuration happens at the reactive center. Strong nucleophiles are required. Good leaving groups matter. Steric hindrance kills the reaction.
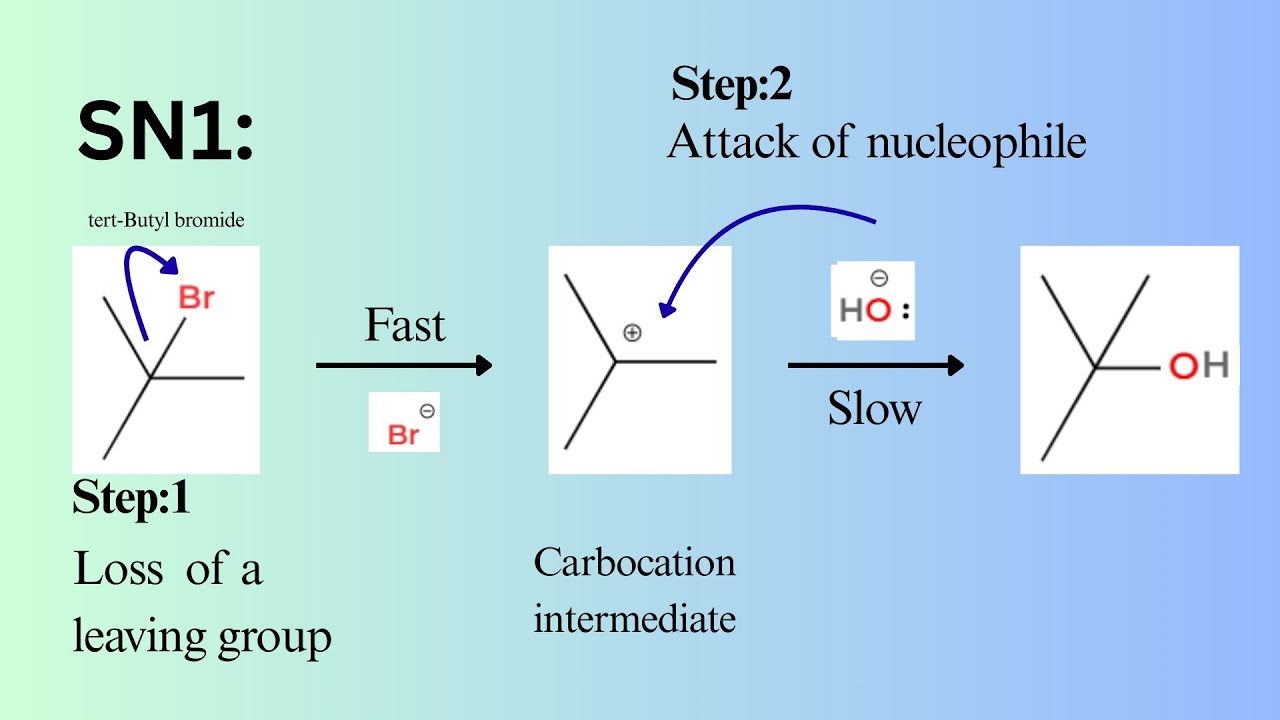
The SN1 mechanism happens in two distinct steps. The leaving group departs first, forming a carbocation intermediate. Then the nucleophile attacks. The rate depends only on the substrate concentration, expressed as rate = k[substrate]. The nucleophile strength does not affect the rate at all. Racemization is common because the carbocation is planar and the nucleophile can attack from either face. Weak nucleophiles are fine. Solvent stability of the carbocation becomes critical. Here is a detail that is not always emphasized in textbooks. Methyl substrates cannot do SN1 because a methyl carbocation is far too unstable. They also cannot do SN1 even under forcing conditions. If you see a primary or methyl substrate and the product shows loss of stereochemistry or rearrangement, something else is happening, possibly an E2 elimination or a radical pathway, not SN1. Solvent choice is where many people make mistakes. Polar aprotic solvents like acetone, DMSO, and DMF accelerate SN2 reactions by leaving the nucleophile relatively unsolvated and more reactive. These solvents are bad for SN1 because they do not stabilize carbocations well. Polar protic solvents like water, methanol, and acetic acid stabilize carbocations through hydrogen bonding and solvation, which accelerates SN1. The same protic solvents slow SN2 by surrounding and the nucleophile.
I learned this the hard way during a graduate lab run. I was trying to convert a secondary tosylate to a methyl ether using sodium methoxide in methanol. I expected clean SN2 with inversion. Instead, I got a mixture that was roughly 60 percent SN1 product with partial racemization and 40 percent elimination. The problem was the solvent. Methanol is protic, which stabilized the carbocation enough to open the SN1 channel even though I had a strong nucleophile present. The workaround was switching to anhydrous DMF with sodium methoxide. The product shifted dramatically toward SN2 inversion. Yield improved from about 35 percent to roughly 72 percent based on gas chromatography. It took me two full days of troubleshooting before I figured out the solvent was the culprit. Nucleophile strength is another area where convention gets oversimplified. A strong nucleophile favors SN2, but if the substrate is tertiary, no nucleophile strength will force SN2. The steric barrier is simply too high. Conversely, a weak nucleophile like water or alcohol can still give fast SN1 on a tertiary substrate because the rate-determining step does not involve the nucleophile at all. Leaving group ability matters for both mechanisms but in slightly different ways. Iodide is generally better than bromide, which is better than chloride, which is better than fluoride. Tosylate and mesylate are excellent leaving groups in both pathways. The difference is that in SN2, a better leaving group primarily lowers the activation energy of the single transition state. In SN1, a better leaving group directly accelerates the rate-determining ionization step. This is why sulfonyl esters are so commonly used in synthetic work.
Get the Full Details
Temperature is a factor that beginners often ignore. Higher temperatures generally favor elimination over substitution in both SN1 and SN2 contexts. If you need substitution, keep the temperature moderate. I usually run SN2 reactions at room temperature or slightly below when possible. SN1 reactions often need gentle warming to promote ionization, but once the carbocation forms, raising the temperature further just increases the elimination side products. There is a counter-intuitive case that causes problems. Neopentyl substrates, which are primary carbons attached to a quaternary carbon, are essentially unreactive in SN2 despite being primary. The steric bulk right next to the reactive center blocks the backside approach. I have seen this come up in process chemistry when someone assumed a primary halide would react readily. Switching to an SN1 condition does not help either because the resulting primary carbocation is too unstable. The practical solution is to use a different synthetic strategy altogether, such as converting the alcohol to a better leaving group and using a different displacement approach, or going through an allylic intermediate if the structure allows it. Another issue is neighboring group participation. Certain substrates with nearby participating groups like sulfur or nitrogen can undergo substitution with complete retention of configuration, even on secondary carbons. This looks like SN2 on the surface but is actually a two-step process where the neighboring group forms a cyclic intermediate and then the nucleophile opens it. If you are confused about why your stereochemistry is not inverting as expected, this is the first thing to check in the structure.
Concentration effects also play a role that is rarely discussed in introductory courses. In SN2 reactions, higher nucleophile concentration directly increases the rate. In SN1 reactions, increasing nucleophile concentration has no effect on the rate of the first step, though it does affect the product distribution once the carbocation forms. This means you can sometimes push a borderline secondary substrate toward SN2 simply by using a high concentration of a strong nucleophile in an aprotic solvent. The biggest practical limitation of SN1 is carbocation rearrangement. If a hydride or alkyl shift can produce a more stable carbocation, it will happen, often rapidly. This means the product you isolate may not reflect the structure of your starting material. I encountered this with a secondary bromide that rearranged to a tertiary carbocation before the nucleophile could attack, giving a product with the nucleophile attached to a different carbon than expected. The only workaround was to avoid SN1 conditions entirely and force the SN2 pathway through solvent and nucleophile selection. Steric effects in SN2 are not always obvious from the substrate alone. Bulky nucleophiles like tert-butoxide or lithium diisopropylamide will favor elimination over substitution even on primary substrates. This is why strong, bulky bases are classified as bases rather than nucleophiles in synthetic planning. If you need substitution on a hindered substrate, smaller nucleophiles like cyanide, azide, or thiolate are much more reliable.
For secondary substrates specifically, the decision between SN1 and SN2 often comes down to a three-way check. First, identify the solvent. Aprotic favors SN2. Protic favors SN1. Second, assess the nucleophile. Strong and concentrated favors SN2. Weak and neutral favors SN1. Third, look at temperature. Lower temperatures favor substitution over elimination in both cases. If all three point the same way, the prediction is usually reliable. When they conflict, as they did in my methanol experiment above, you need to run a small test reaction before committing to the full scale. One more practical note on workup. SN1 reactions in protic solvents often produce acidic byproducts from the leaving group, such as HBr or HCl. This can cause product decomposition if the product is acid-sensitive. Quenching with a mild base like sodium bicarbonate during workup is standard practice. SN2 reactions in aprotic solvents generally produce less acidic waste, but the workup still needs to remove metal salts and excess nucleophile. Ion-exchange resins or careful aqueous extraction are usually sufficient.