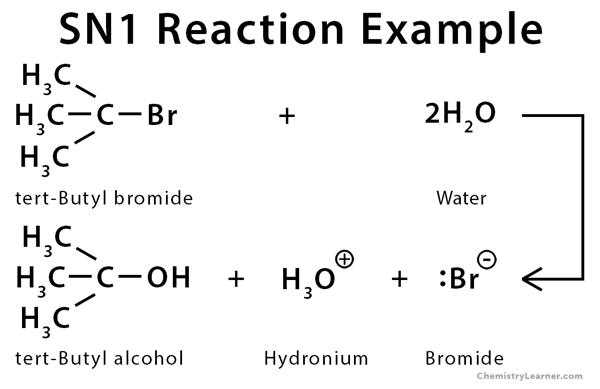
The Mechanism Works Differently Than Your Textbook Suggests
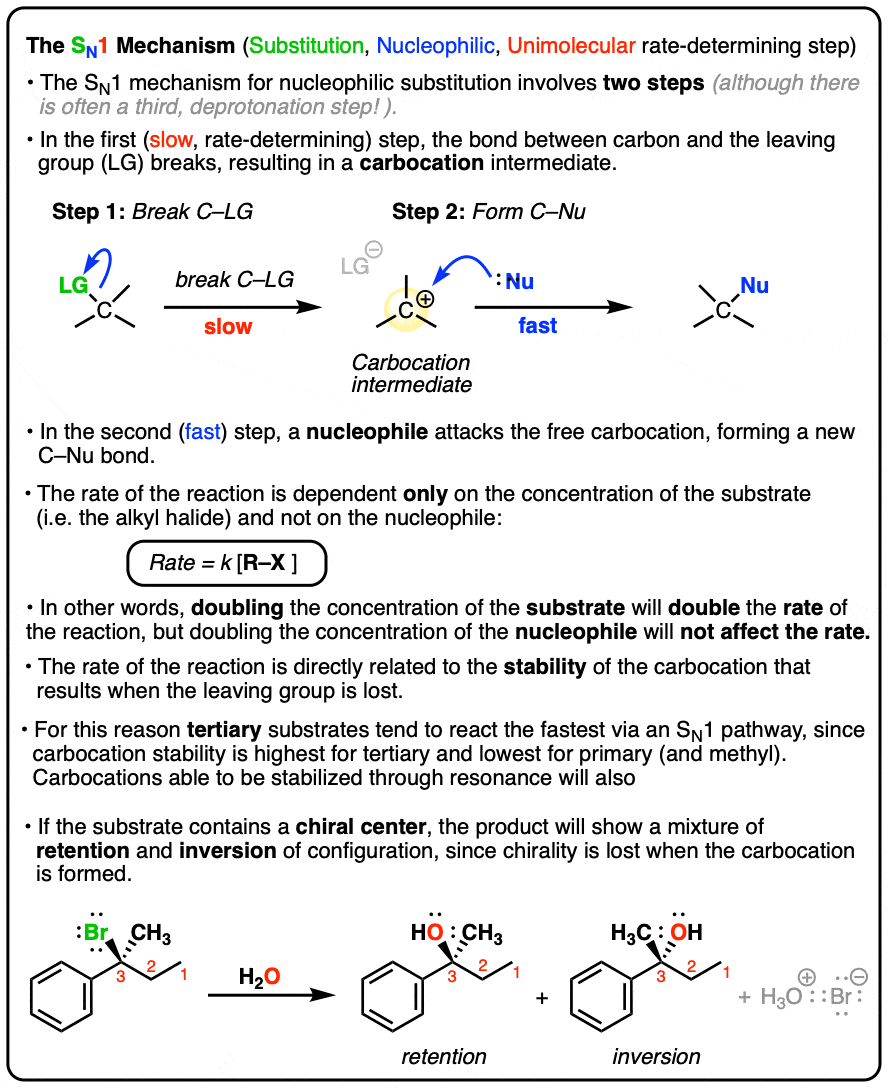
The SN1 reaction is fundamentally a two-step process where the leaving group departs first, forming a carbocation intermediate, and then the nucleophile attacks. That order matters more than people realize. In practice, the rate-determining step is the formation of that carbocation, which means the concentration of the nucleophile has zero effect on how fast the reaction proceeds. Most students miss this when they're studying for exams. I spent a solid week troubleshooting a reaction back in grad school where I thought I had an SN1 pathway going. The substrate was a secondary benzylic halide, and I was expecting clean substitution in ethanol. Instead, I got a mixture of substitution and elimination products along with some rearranged material. The problem was that the carbocation intermediate was long-lived enough to undergo a hydride shift before the nucleophile could capture it. I ended up running the reaction at lower temperature and using a more nucleophilic solvent, which shortened the lifetime of that intermediate and gave me the product I actually wanted. The yield went from roughly 40% to about 78% once I stopped fighting the rearrangement. The biggest misconception about SN1 reactions is that they only happen with tertiary substrates. That's half true. Tertiary carbons form stable carbocations, yes, but secondary substrates can and do undergo SN1 reactions regularly when the resulting carbocation gets resonance stabilization. Benzylic and allylic systems are the classic examples here. A secondary benzylic chloride in aqueous ethanol will go SN1 comfortably at room temperature, while a simple secondary alkyl chloride will sit there and do nothing under identical conditions.
Common Sn1 Reaction Examples and What Actually Happens
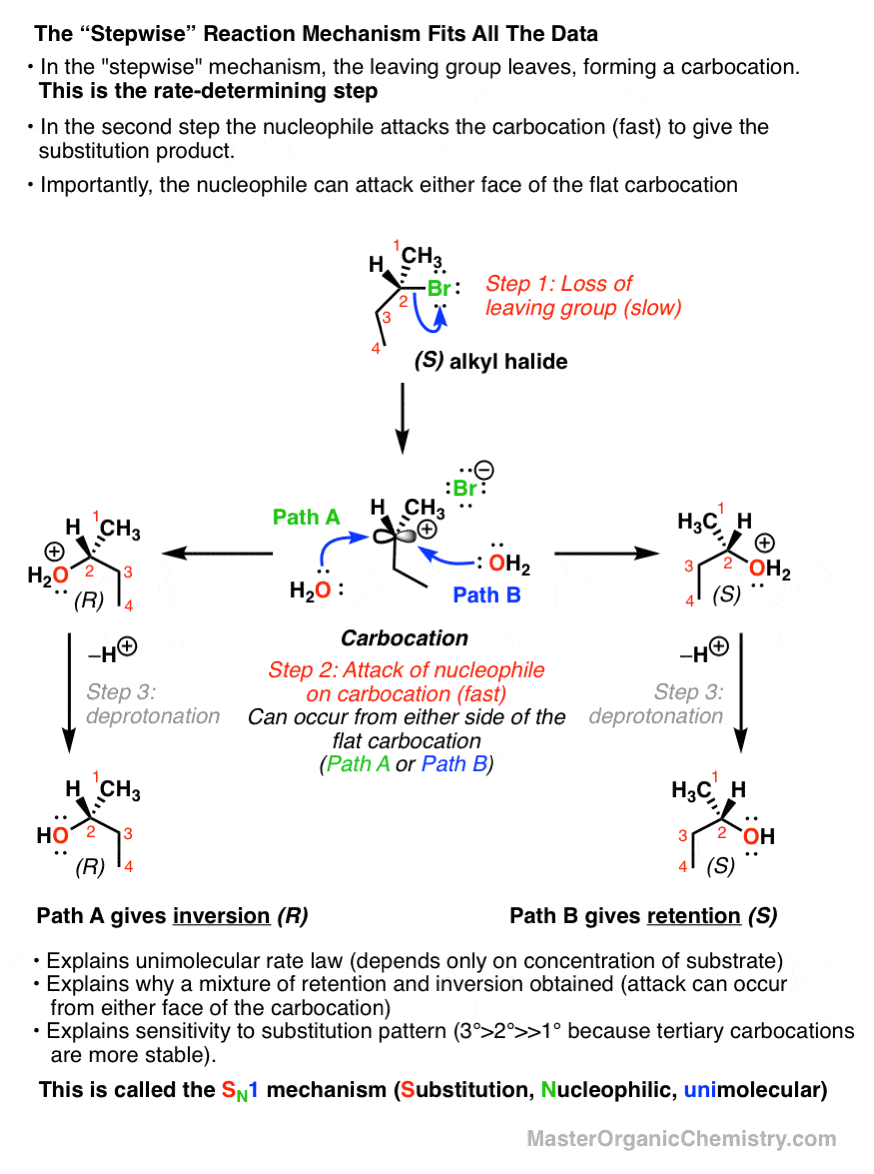
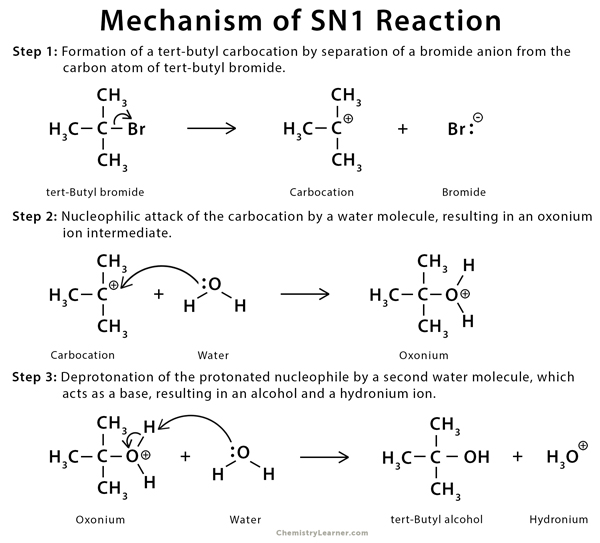
tert-Butyl bromide with water: This is the textbook example everyone learns first. The bromide leaves, forming a tert-butyl carbocation, and water attacks to give tert-butyl alcohol. The reaction is clean because the tertiary carbocation is highly stable and there's no possibility of rearrangement improving the structure. It proceeds readily in a polar protic solvent without any external catalysis. Benzyl chloride with methanol: The benzyl carbocation is stabilized by resonance with the aromatic ring. Methanol acts as both solvent and nucleophile here, producing benzyl methyl ether. The reaction goes faster than you'd expect for a primary-like substrate because that resonance stabilization lowers the activation energy for the ionization step significantly. 1-Phenylethyl chloride in aqueous acetone: This secondary benzylic substrate undergoes solvolysis to give a mixture of the alcohol and the corresponding ether if ethanol is present alongside water. The carbocation intermediate is planar, so you get racemization at the stereocenter. If you start with an enantiomerically pure sample, you'll end up with a racemic product. This isn't a flaw in the reaction—it's just how the mechanism works, and it catches people off guard when they're trying to preserve stereochemistry.
Cyclohexyl bromide with formic acid: Formic acid is a highly ionizing solvent that promotes SN1 reactions even on secondary substrates that wouldn't react in less polar media. The bromide leaves, the formate anion attacks the carbocation, and you get cyclohexyl formate. This example is useful because it shows how solvent choice can override the usual substrate restrictions. Here's something most sources don't emphasize enough: the leaving group ability in SN1 reactions follows a different priority than SN2. In SN2, steric factors dominate. In SN1, it's almost entirely about how well the leaving group can stabilize the negative charge as it departs. Iodide is an excellent leaving group in both mechanisms, but the difference between chloride and tosylate matters enormously in SN1. A chloride on a tertiary carbon might sit inert for hours at room temperature, while the same carbon with a tosylate group will ionize within minutes in the right solvent. The bond strength of C-Cl is simply too high for easy heterolytic cleavage without additional stabilization. Carbocation rearrangement is the silent product killer in SN1 chemistry. A 1,2-hydride shift or 1,2-methyl shift can occur if it leads to a more stable carbocation, and this happens fast—often faster than the nucleophile can attack. I once ran a reaction expecting a straightforward substitution on a secondary carbon adjacent to a quaternary center, and the major product came from a methyl shift that converted the secondary carbocation into a tertiary one. The expected product was maybe 15% of the mixture. I learned to map out every possible rearrangement pathway before running these reactions instead of assuming the direct substitution product would dominate.
Get the Full Details

The stereochemical outcome deserves another look. Because the carbocation is sp2 hybridized and planar, nucleophilic attack can occur from either face. In ideal conditions, you get complete racemization. In practice, the leaving group often partially blocks one face of the carbocation as it departs, leading to slight excess of inversion over retention—usually around 55 to 60% inversion. This is called partial racemization with some inversion, and it's the reason you can't use SN1 reactions when you need to preserve optical purity. Another practical limitation: SN1 reactions struggle with primary substrates except in special cases. A primary carbocation is too unstable to form under normal conditions, so primary halides almost exclusively undergo SN2. The exception is when the primary carbon is benzylic or allylic, where resonance provides enough stabilization. Even then, you'll often see a competing SN2 pathway unless the reaction conditions heavily favor ionization. If you're planning a synthesis and need clean substitution without rearrangement or racemization concerns, consider switching to an SN2 protocol instead. Using a strong nucleophile in an aprotic solvent like DMF or acetonitrile with a primary or secondary substrate will give you predictable inversion of configuration and no carbocation intermediates to rearrange. The SN1 pathway is useful when your substrate simply can't undergo SN2—tertiary centers, for instance—but it comes with the tradeoffs of potential rearrangement and loss of stereochemical control.
The solvent effect is perhaps the most practically important variable. Polar protic solvents like water, methanol, ethanol, and acetic acid stabilize the carbocation intermediate and the leaving group anion through hydrogen bonding and dipole interactions. This dual stabilization dramatically lowers the activation energy for the ionization step. Switching from a 50:50 ethanol-water mixture to pure water can increase the reaction rate by an order of magnitude for the same substrate. Nonpolar solvents essentially shut down SN1 reactions because they can't stabilize the charged intermediates.
Practical Considerations Before Running an SN1 Reaction
Temperature control matters more than in SN2 reactions because the carbocation intermediate is higher in energy and more reactive. Running the reaction too warm increases the chance of elimination side products and rearrangement. Keeping it cool gives the nucleophile a better shot at capturing the intermediate before it does something unwanted. I typically run SN1 solvolyses at or slightly below room temperature unless the substrate is particularly unreactive. Concentration of the nucleophile doesn't affect the rate, but it does affect the product distribution when competing nucleophiles are present. In a mixed solvent system like water-ethanol, you'll get both the alcohol and the ether product, and the ratio depends on the relative concentrations and nucleophilicity of water versus ethanol. Water is a better nucleophile in this context despite being smaller because it's present at much higher effective concentration in aqueous solutions. Tracking these reactions can be tricky because the rate depends on substrate ionization, which isn't always obvious visually. I've used conductivity measurements to monitor carbocation formation in real time—the solution becomes more conductive as ions are generated. For routine work, thin-layer chromatography is sufficient, but keep in mind that carbocation intermediates can continue to react on the TLC plate if they're long-lived enough, giving you spots that don't correspond to your intended product.