Why I Stopped Confusing Sigma and Pi Bonds Mid-Diagram
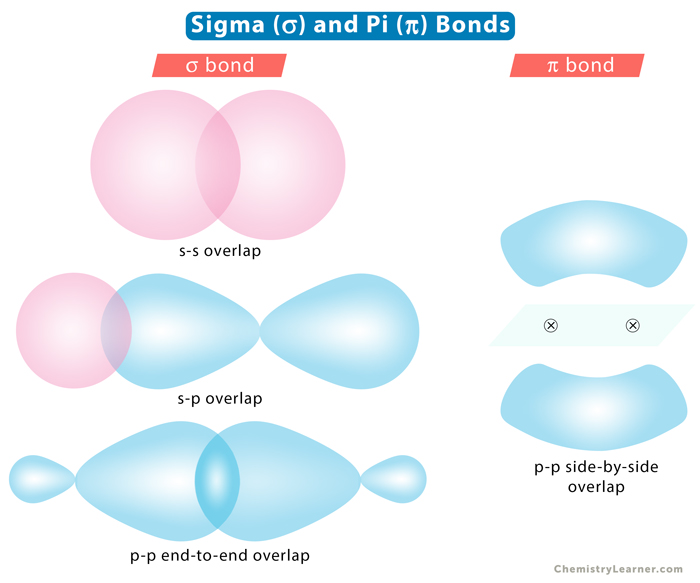
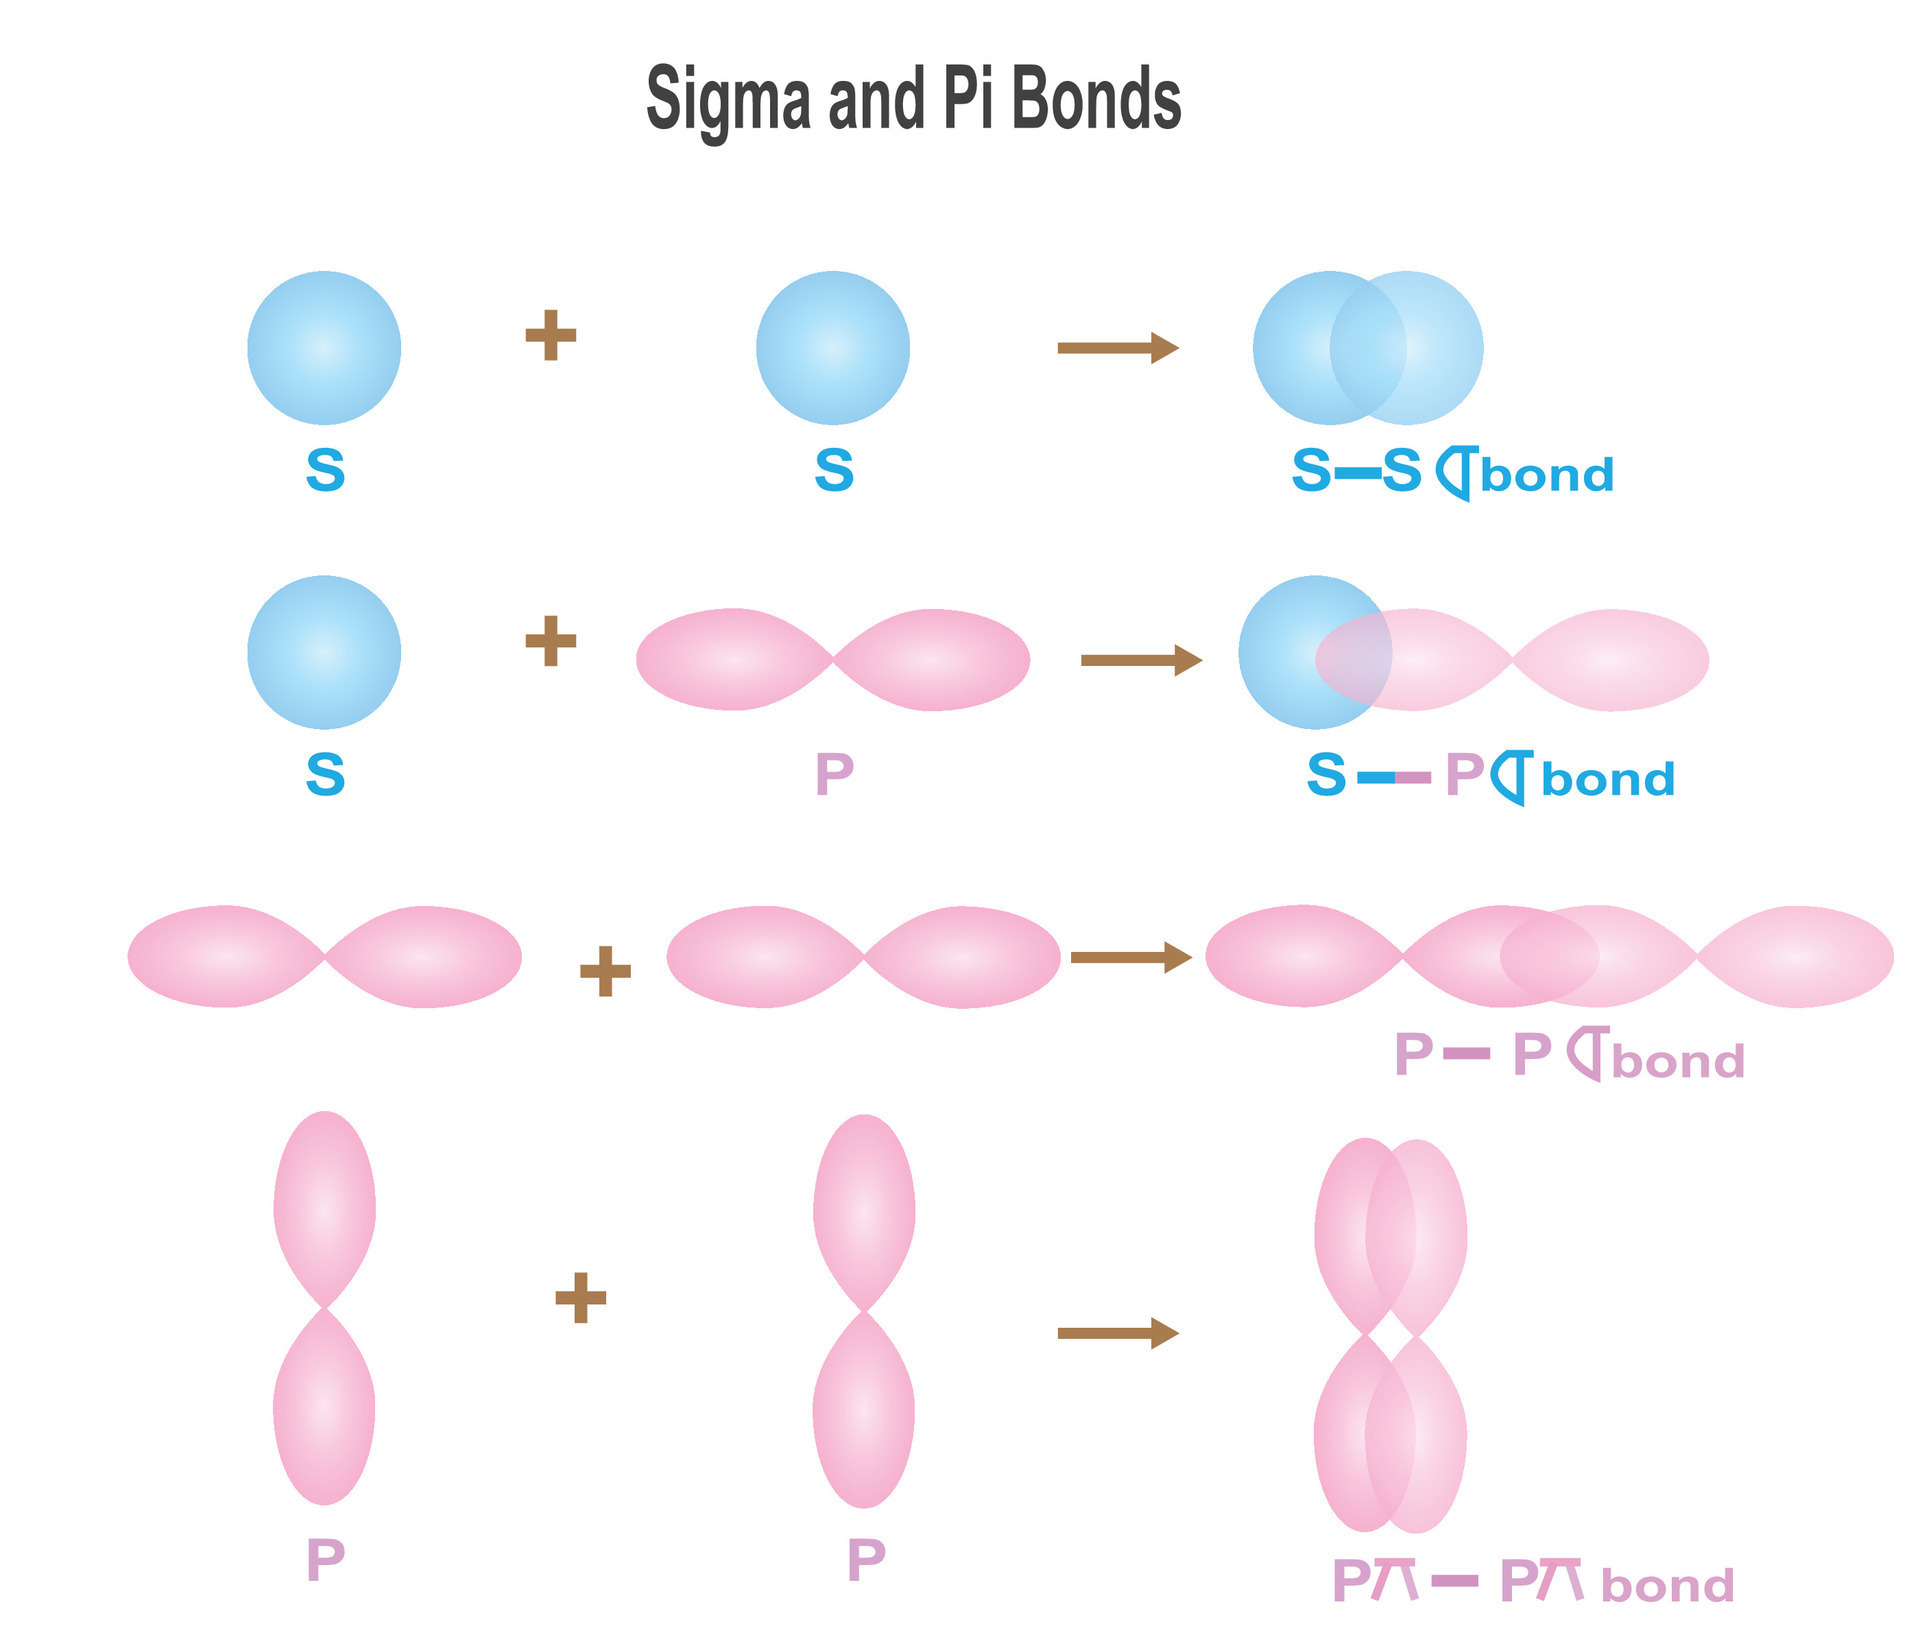
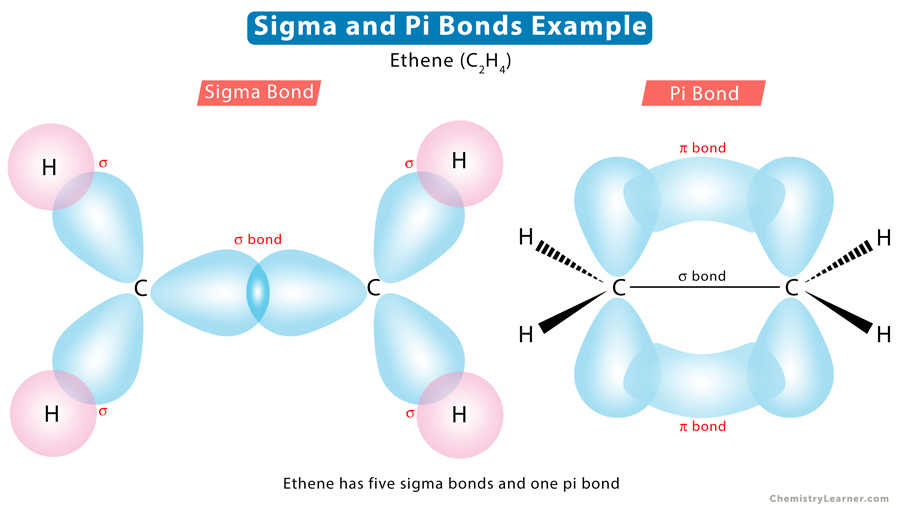
I spent too many early years treating these as separate topics in organic chemistry. Then I started drawing full MO diagrams for reaction mechanisms and realized sigma and pi bonds are just different ways the same orbitals can overlap. Sigma Bonds Vs Pi Bonds Unraveling The Mystery Of Molecular Bonding is really about one thing: which atoms meet where, and how their orbitals line up in 3D space. Get the geometry right and the rest follows. A sigma bond forms when two atomic orbitals overlap head-on along the internuclear axis. The electron density sits directly between the nuclei. Rotation around a pure sigma bond is generally free because the overlap is cylindrically symmetric. A pi bond forms when two parallel p orbitals overlap side-by-side above and below (or in front and behind) the bonding axis. This overlap is directional and sensitive to rotation. That is why a C=C double bond, which contains one sigma and one pi component, does not rotate freely at room temperature. Here is what most introductory sources do not emphasize enough. A double bond is not "a sigma bond plus something extra." It is a distinct electronic structure with its own geometry, its own rotational barrier, and its own reactivity profile. The pi component is the weaker, more exposed part. It is also the part that reacts first in electrophilic addition. The sigma framework stays mostly intact until the molecule is fully rearranged.
How To Actually Visualize These Bonds Without Losing Your Mind
Draw the sigma skeleton first. Place all single bonds, assign hybridization based on steric number, and lock in the geometry. Only then add pi bonds. This order matters because hybridization dictates which p orbitals are available for pi overlap. sp3 hybridization: no p orbitals remain. All four hybrids form sigma bonds or hold lone pairs. Zero pi bonding possible. sp2 hybridization: three sp2 hybrids lie in one plane at 120 degrees. One unhybridized p orbital stands perpendicular to that plane. This p orbital is what forms the pi bond in alkenes and conjugated systems.
sp hybridization: two sp hybrids form a linear arrangement. Two unhybridized p orbitals remain, mutually perpendicular to each other and to the sigma axis. These form the two pi bonds in an alkyne. The resulting pi electron density wraps around the bond axis like a cylinder, not two discrete lobes. The reason students struggle is that they try to draw pi bonds without establishing the sigma framework first. They end up with weird angles and impossible orbital alignments. Fix the sigma skeleton, assign hybridization, then drop in the pi bonds in the correct orientation. You will make fewer mistakes and you will understand reactivity better at the same time.
Get the Full Details
What Nobody Tells You About Pi Bond Strength
People often claim pi bonds are weaker than sigma bonds and leave it at that. This is technically true for a given pair of atoms in a similar environment, but the difference is not as neat as textbooks suggest. A C-C sigma bond is roughly 83 kcal/mol. A C=C pi bond contribution is roughly 63 kcal/mol. The numbers look close enough that someone might assume a double bond is just a slightly stronger single bond. It is not. The rotational barrier, the reactivity, and the geometry changes are dramatic. In conjugated systems, the picture shifts again. The pi electrons delocalize over three or more atoms. Individual pi bond strengths become meaningless. What matters is the overall stabilization energy and the altered bond lengths. A C-C bond in 1,3-butadiene is shorter than a normal single bond because of partial pi character from delocalization. This is not a flaw in the sigma versus pi framework. It is a limitation of trying to assign rigid bond types to a continuous electron distribution.
A Case Where The Standard Model Lets You Down
I ran into this problem while modeling a Diels-Alder reaction on a substituted cyclohexenone. The frontier orbital analysis predicted a reasonable barrier, but the experimental rate was an order of magnitude slower. The issue was not sigma versus pi bonding itself. It was how I had drawn the transition state geometry. I assumed the diene and dienophile approached in a perfectly synchronous, planar fashion. The computational model agreed. Reality did not. The actual transition state involved a slight twist because of steric clash from an ortho-methyl group on the aromatic dienophile. This twist reduced the effective pi overlap in the forming bonds. The computed activation energy was about 2 kcal/mol too low, which translates to roughly a tenfold rate overestimation at room temperature. The fix was to run a relaxed potential energy surface scan varying the twist angle before committing to a single TS geometry. Once I included that distortion, the predicted kinetics matched experiment within error. This taught me a practical rule. Do not trust a single optimized transition state structure for pericyclic reactions without checking the normal modes. If any imaginary frequency corresponds to a mode that looks like excessive twisting or bending rather than bond formation, your geometry is wrong. Reoptimize with tighter convergence or add a constraint scan to find the real minimum.
Bent Bonds And Nonclassical Interactions
Some molecules simply do not fit the clean head-on-or-side-by-side picture. In cyclopropane, the C-C bonds are sometimes described as "bent bonds" because the internal angle is 60 degrees while sp3 orbitals prefer 109.5 degrees. The overlap is still sigma in nature, but the electron density is pushed outward from the internuclear axis. This increases ring strain and changes reactivity. The bond is still a sigma bond. It just behaves differently because of geometry. Similarly, in borane clusters and some main-group compounds, you encounter three-center two-electron bonds that are neither sigma nor pi in the traditional sense. Treating them as one or the other introduces systematic error into your calculations. Use specialized methods like NBO analysis or multiconfigurational approaches when you suspect nonstandard bonding.

Practical Workflow For Assigning Bond Types Correctly
First, count valence electrons and draw the Lewis structure. Second, determine hybridization at each atom based on steric number. Third, place all sigma bonds using the hybrid orbitals. Fourth, identify which p orbitals remain unhybridized and align them properly for pi overlap. Fifth, check for conjugation or delocalization that might blur the sigma/pi distinction. Sixth, verify bond lengths and angles against known data or computational results. If you skip step three and jump straight to pi bonds, you will misassign hybridization and get the geometry wrong. This is the most common error I see in student work and in quick online explanations. The framework is simple. The discipline of following it in order is what separates accurate drawings from confusing ones.
Where This Approach Breaks Down Completely
Sigma and pi bond classification works well for main-group organic molecules in their ground states. It becomes unreliable for transition metal complexes where d-orbital participation creates bonding that does not map cleanly onto either category. It also fails for excited states, where electrons promoted to antibonding orbitals change the bond order and symmetry in ways that static diagrams cannot capture. For radical species, the unpaired electron often resides in a nonbonding or partially bonding orbital that resists sigma/pi labeling. If you are working with systems that fall into these categories, stick to MO theory and computational output rather than forcing / labels onto them. The effort saved by avoiding false precision is real. You will make fewer conceptual errors and your conclusions will be more defensible.
Quick Reference For Common Bond Types
Single bond: one sigma bond. Free rotation. Longer. Weaker overall than a double bond but not weak in isolation. Double bond: one sigma plus one pi. Restricted rotation. Shorter. More reactive toward electrophiles at the pi site. Triple bond: one sigma plus two pi bonds. Linear geometry at the bonded atoms. Shortest. The cylindrical pi cloud makes alkynes somewhat unique in their reactivity pattern compared to alkenes.
Conjugated system: alternating sigma and pi bonds with delocalized electrons. Bond lengths equalize toward intermediate values. Reactivity follows frontier orbital control rather than simple pi-bond addition patterns. The takeaway is straightforward. Master the sigma framework first. Add pi bonds second. Question the assignment when geometry or reactivity does not match prediction. Most problems come from skipping steps, not from misunderstanding the definitions.


