Orbital Overlap Is Not What Your Professor Made It Sound Like
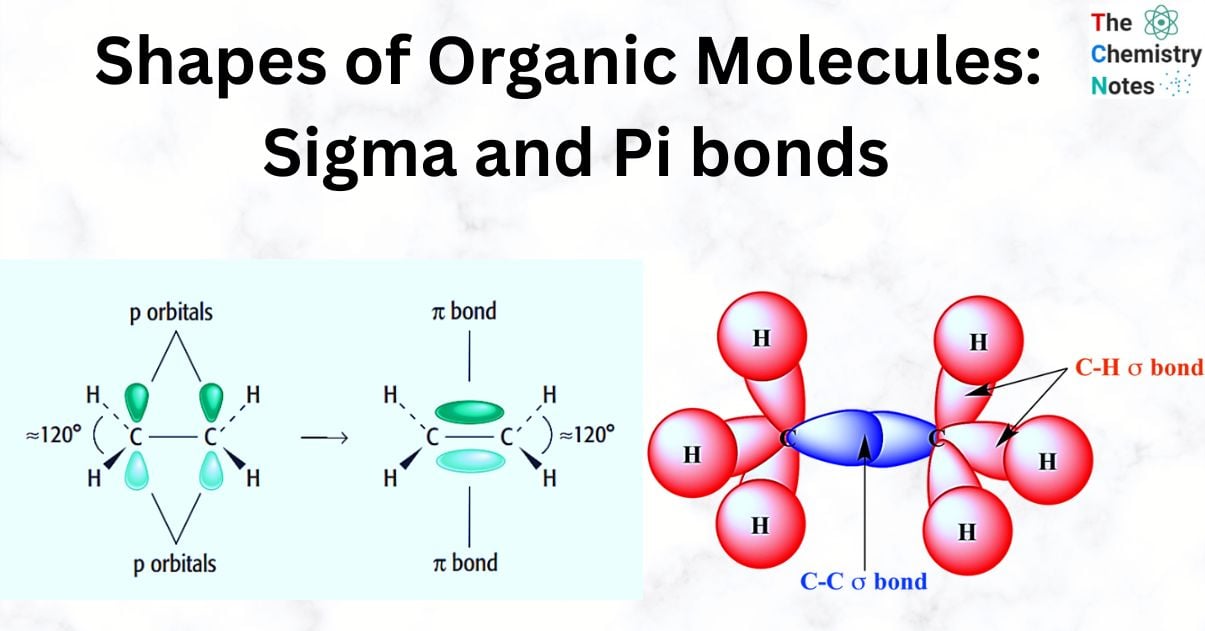
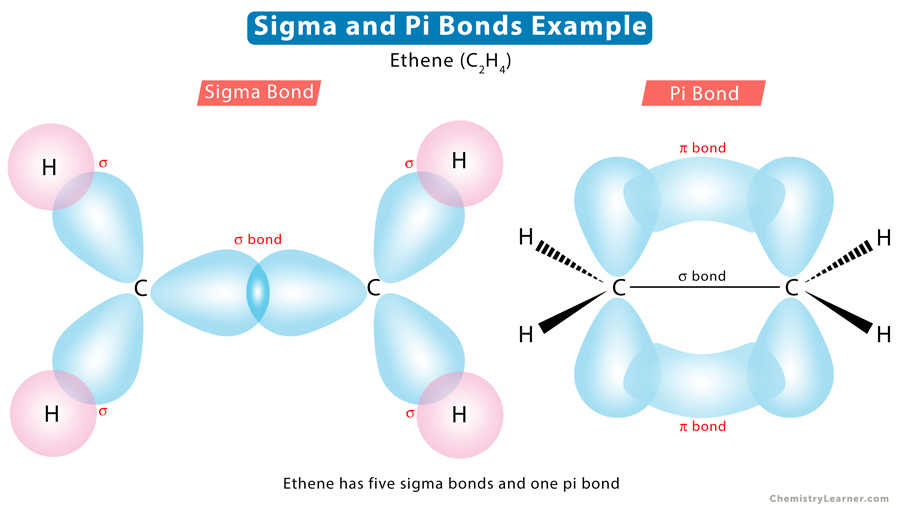
Sigma bonds and pi bonds aren't mystical categories. They're just two ways atomic orbitals can overlap when two atoms get close enough to share electrons. That's it. The rest of the confusion comes from people treating them like separate concepts instead of describing the same event from different geometric angles. A sigma bond forms when orbitals overlap along the internuclear axis—the direct line between two nuclei. s-s overlap, s-p overlap, p-p head-on overlap, hybrid-hybrid overlap. All of these produce sigma bonds. The electron density is concentrated between the nuclei, and rotation around that bond axis is essentially free at room temperature because the overlap doesn't change as you twist. A pi bond forms when unhybridized p orbitals sit parallel to each other and overlap sideways, above and below the plane of the internuclear axis. You need two parallel p orbitals for this to happen, which means one atom must still have an unhybridized p orbital available after sigma bonding has taken place. That's why you see pi bonds exclusively in double and triple bonds, never in single bonds. Single bonds are always pure sigma. The pi component is what turns a single bond into a double bond or triple bond.
The weakness of pi bonds relative to sigma bonds is what makes organic chemistry interesting. A typical C-C sigma bond holds at roughly 347 kJ/mol. A C=C double bond isn't twice that—it's about 614 kJ/mol. The pi component contributes only about 267 kJ/mol, significantly weaker than the sigma bond it rides on. This matters because when molecules react, the pi electrons are the ones getting attacked first. They're higher in energy, more exposed, and sitting further from the nucleus where they're held less tightly. Nucleophiles and electrophiles find pi bonds long before they ever notice the sigma framework. I spent a semester trying to predict the reactivity of a substituted cyclohexenone system, and I kept getting the regiochemistry wrong because I was thinking about the pi bond as if it were just a second bond tacked onto a sigma bond. The actual issue was that the carbonyl pi bond and the alkene pi bond were conjugated—delocalized across three atoms. The electrons weren't sitting in two separate pi bonds at all. They existed as a single molecular orbital spread across the C-C-C-O framework. Once I started drawing the resonance structures properly and tracking where the HOMO electron density actually lived, the reaction made sense. The nucleophile went to the beta carbon, not the carbonyl carbon, because that's where the LUMO coefficient was largest. This is the kind of thing that doesn't get emphasized enough in introductory courses.
Hybridization Dictates What Bonds Are Even Possible
The geometry of a molecule is determined by hybridization, and hybridization determines whether pi bonds can form at all. An sp3 carbon has four equivalent hybrid orbitals arranged tetrahedrally. All four are used for sigma bonding. There are no leftover p orbitals. No pi bonds possible. Methane, ethane, cyclohexane—these molecules are purely sigma frameworks. An sp2 carbon uses three hybrid orbitals for sigma bonding in a trigonal planar arrangement and keeps one unhybridized p orbital perpendicular to that plane. That p orbital is what forms the pi bond in alkenes. Ethylene is the simplest example—each carbon is sp2, the C-C sigma bond comes from sp2-sp2 overlap, and the pi bond comes from the two remaining p orbitals overlapping sideways. The bond is rigid. You cannot rotate around a double bond without breaking the pi overlap, and that requires roughly 260 kJ/mol. At room temperature, that barrier is insurmountable, which is why cis and trans isomers are stable, isolable compounds. An sp carbon keeps two unhybridized p orbitals. It can form two pi bonds using both, giving you a triple bond. Acetylene is the textbook case—sp-sp sigma bond down the middle, two pi bonds arranged perpendicularly around it, one from p-p overlap in one plane and the other from p-p overlap in the perpendicular plane. The molecule is linear because sp hybridization produces a 180-degree angle between the two sigma bonds.
Get the Full Details
Conjugation Changes Everything
When pi bonds are separated by exactly one sigma bond, they conjugate. The p orbitals on adjacent atoms overlap not just with their immediate neighbor but across the entire conjugated system, forming a set of delocalized molecular orbitals. Butadiene is the simplest case—four carbons, three sigma bonds in the chain, two pi bonds that merge into a single delocalized pi system with three molecular orbitals: one bonding, one nonbonding, and one antibonding. The six pi electrons fill the bonding and nonbonding orbitals. This delocalization provides extra stability—the resonance energy of butadiene is roughly 15 kJ/mol compared to two isolated double bonds. It also changes reactivity dramatically. Conjugated systems undergo 1,4-addition instead of 1,2-addition because the electrophile attacks the terminal carbon of the system, not the carbon directly involved in the pi bond. The intermediate is an allylic carbocation, which is stabilized by the adjacent pi system. This is foundational for understanding Diels-Alder reactions, aromatic substitution, and practically every reaction in advanced organic chemistry. I had a grad student once who couldn't figure out why a particular conjugated enone wasn't reacting under standard Grignard conditions. The problem wasn't the reagents or the temperature. It was that the conjugated system was extended into an aromatic ring on one side, which lowered the LUMO energy enough that the Grignard reagent was performing 1,4-addition instead of the expected 1,2-addition to the carbonyl. We spent two days checking spectra before someone pointed out the extended conjugation. The workaround was switching to a cuprate reagent, which is softer and more selective for 1,4-addition anyway. This is one of those practical realities you only learn by making the same mistake.
Computational Chemistry Has blind Spots With Pi Systems
If you're modeling molecules with pi bonds using standard DFT functionals, be aware that many of them struggle with dispersion interactions and charge transfer in conjugated systems. B3LYP, which was the workhorse for years, systematically underestimates barrier heights for reactions involving pi bond breaking and over-stabilizes delocalized transition states. For accurate pi-system calculations, you're better off using a functional like B97X-D or M06-2X, which include empirical dispersion corrections and are parameterized for noncovalent interactions. The downside is computational cost.B97X-D with a reasonable basis set like 6-311+G(d,p) can take roughly three to five times longer than B3LYP for the same system. For small molecules under 30 atoms, this is manageable. For larger conjugated systems or proteins with pi-stacking interactions, it becomes a genuine constraint. If you're working with systems over 100 atoms and need pi-bond accuracy, semi-empirical methods like GFN2-xTB offer a reasonable compromise at a fraction of the cost, though they sacrifice quantitative precision for speed.
Bond Lengths Reveal More Than You Think
Looking at bond lengths is one of the most underutilized tools for diagnosing the character of bonds in a molecule. A standard C-C single bond is 1.54 angstroms. A C=C double bond is 1.34 angstroms. But in conjugated systems, these values shift. In benzene, all six C-C bonds are identical at 1.39 angstroms—intermediate between single and double because the pi electrons are fully delocalized. In butadiene, the central C-C bond is 1.47 angstroms, shorter than a typical single bond because of partial pi character from conjugation. When I'm analyzing an unknown product from a reaction, the first thing I check after the NMR is the X-ray crystal structure if I have it, or at least reliable computational bond lengths. Deviations from expected values tell you immediately whether conjugation is operating, whether there's steric strain distorting the geometry, or whether a bond has partial ionic character that's shortening or lengthening it unexpectedly. A C=O bond that comes out at 1.23 angstroms instead of the expected 1.21 suggests significant single-bond character from resonance donation, which usually means you're dealing with an amide or an enol rather than a simple ketone.

The Real Limitation Nobody Talks About
Sigma and pi bond theory works beautifully for small, well-defined molecules. It breaks down in several important scenarios. Transition metal complexes are the first problem—the d orbitals participating in bonding don't fit neatly into the sp/sp2/sp3 hybridization model. You get delta bonds, phi bonds, and various degrees of covalent and ionic character that simple orbital overlap diagrams can't capture. For organometallic chemistry, you need crystal field theory or ligand field theory, not sigma-pi bond language. Bonded solids are another area where the model fails. Graphite has pi electrons that are delocalized across an entire two-dimensional sheet, not between two atoms. The concept of a discrete pi bond doesn't apply—you have a pi band in the electronic structure, not individual bonds. Diamond is purely sigma bonded in a three-dimensional network, but calling it a collection of sigma bonds misses the fact that it's a single continuous crystal with band structure, not discrete molecules. Even for organic molecules, highly strained systems like cyclopropane don't behave according to standard sigma bond models. The C-C bonds in cyclopropane are bent, with poor orbital overlap that creates what's called banana bonds or theta bonds. The bond strength is lower than a normal sigma bond, and the electrons are pushed outward, making cyclopropane unusually reactive despite having only single bonds. The Hückel model and simple MO theory don't account for this strain effect.
If you're working with any of these edge cases, you need to move beyond the basic sigma-pi framework. For strained ring systems, consider using computational methods that account for strain energy explicitly. For metals, switch to ligand field theory. For extended pi systems, use band theory or periodic DFT rather than trying to force molecular orbital theory onto a solid-state problem.