The Practical Side of Lewis Acid-Base Chemistry
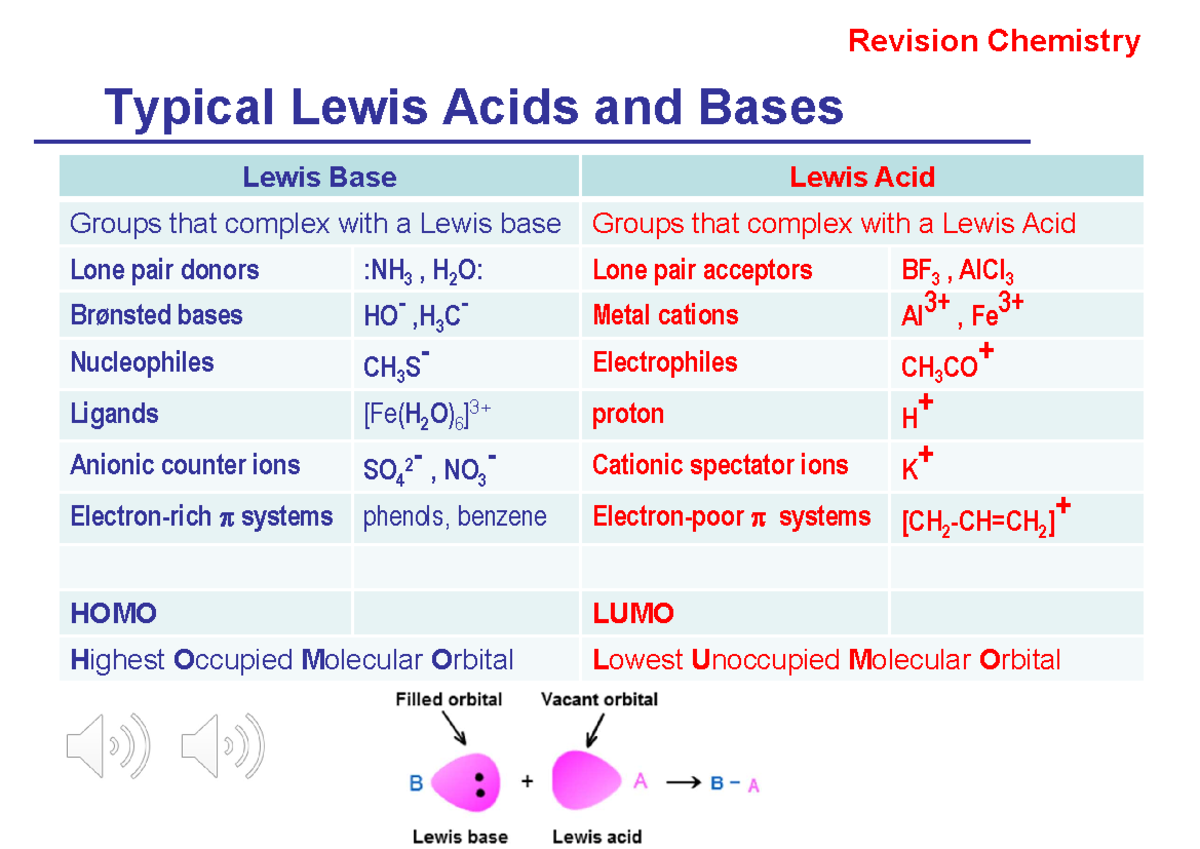
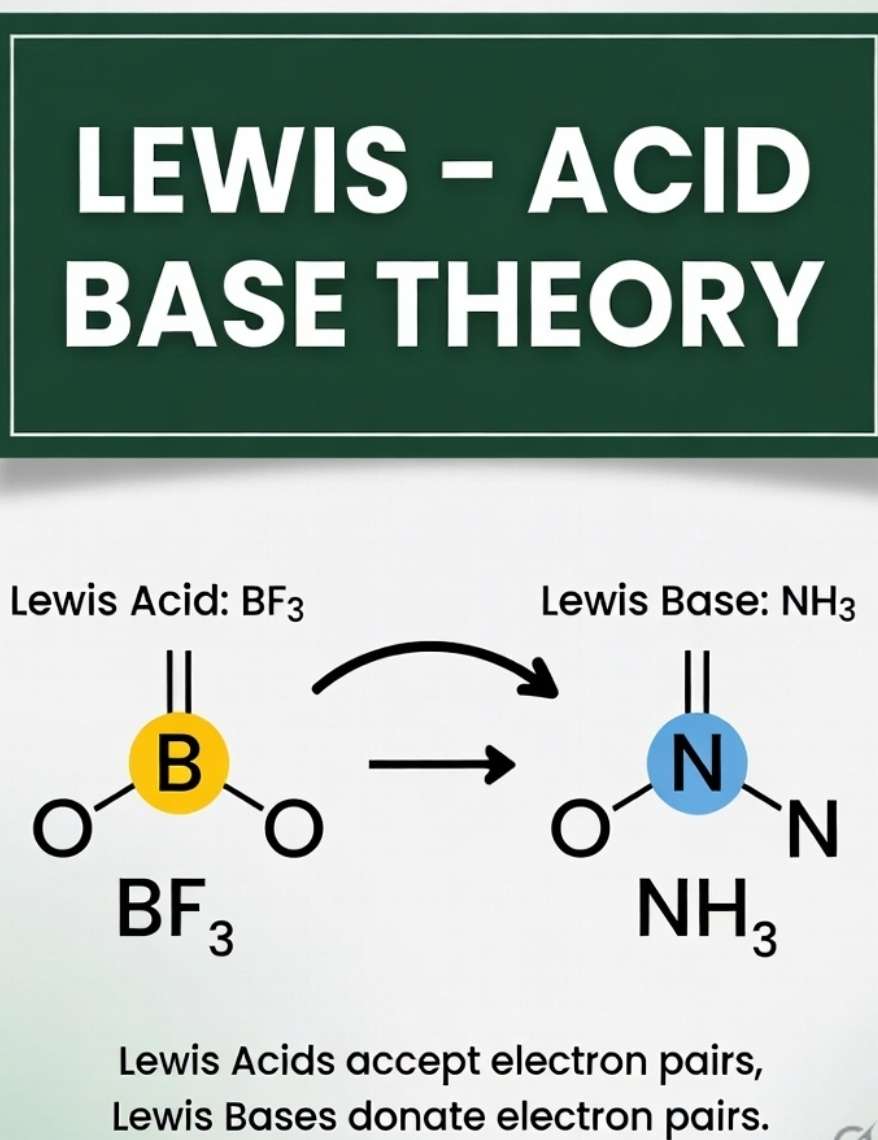
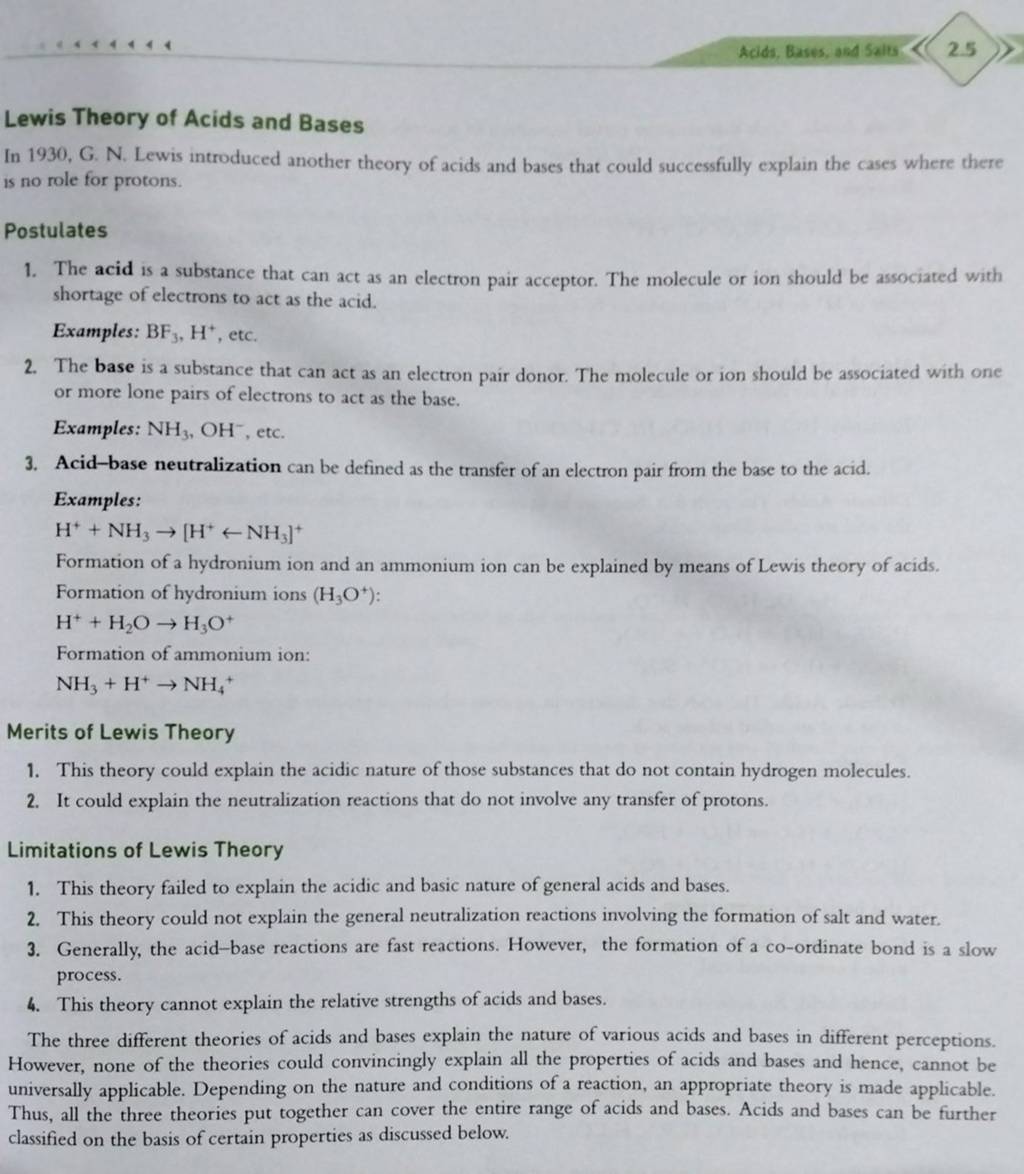
Most textbooks present Lewis acids and bases as simple definitions. An acid accepts an electron pair. A base donates one. That's accurate but completely missing the part that matters when you're actually running a synthesis at 2 AM and something isn't working. The real world of Lewis acid-base chemistry is messier than the textbook version, and the gap between theory and practice is where most people lose time and material. Lewis acid catalysis works by coordinating to a substrate and making it more electrophilic. That coordination lowers the activation energy of a downstream reaction. The classic example is using AlCl3 in a Friedel-Crafts acylation. The aluminum coordinates to the carbonyl oxygen of the acyl chloride, pulling electron density away and making the carbon much more susceptible to attack by an aromatic ring. This is standard undergraduate material. What the undergraduates rarely learn is that the choice of Lewis acid dramatically changes the outcome beyond just "making things more reactive." I spent three days troubleshooting a Diels-Alder reaction that refused to proceed at the recommended temperature. The literature said Zn(OTf)2 should lower the activation barrier sufficiently. Instead, the reaction remained dormant at room temperature and only converted sluggishly at 80 degrees Celsius, which was already near the decomposition point of my diene. The issue turned out to be solvent coordination. My solvent, THF, was competing with the diene for the zinc center. The Lewis acid was busy solvated, not activating my substrate. Switching to dichloromethane, a non-coordinating solvent, restored the expected rate. The reaction went to completion in two hours at room temperature instead of twelve hours at 80 degrees. This kind of solvent-Lewis acid interaction is something you learn the hard way if nobody has told you already.
How Lewis Acids Actually Operate in Practice
The strength of a Lewis acid isn't a single number you look up and apply universally. It depends on the substrate, the solvent, the temperature, and the mechanism. Hard Lewis acids like Mg2+ and Al3+ prefer hard bases like oxygen donors. Soft Lewis acids like Pd2+ and Pt2+ prefer soft bases like phosphines and sulfides. This is the HSAB principle, and it's useful but incomplete. In practice, you'll find that a "hard" acid like BF3 can catalyze reactions with soft nucleophiles if the geometry allows proper orbital overlap, while a supposedly appropriate hard acid might fail completely on a soft substrate due to kinetic barriers that thermodynamic tables don't predict. The most commonly used Lewis acids fall into a few practical categories. Boron-based acids like BF3·OEt2 and B(CH3)3 are workhorses for activating carbonyls and epoxides. Aluminum chlorides and triflates handle Friedel-Crafts reactions and Mukaiyama aldol condensations. Titanium isotope complexes, particularly TiCl4 and Ti(OiPr)4, dominate asymmetric transformations like the Sharpless epoxidation. Tin compounds, especially SnCl4, are valuable when you need a milder alternative to aluminum. Each has specific handling requirements and incompatibilities that matter more than their theoretical reactivity rankings. I learned about tin chloride the hard way. I was running a Mukaiyama aldol using TiCl4 because that's what the procedure specified, but the yield was consistently poor at around forty percent with significant decomposition. The starting silyl enol ether was clearly reacting, but the product kept degrading. I switched to SnCl4 as a milder Lewis acid and the yield jumped to seventy-eight percent. The TiCl4 was simply too aggressive for the functional groups present in my substrate. No amount of temperature adjustment fixed it. This is the kind of detail that doesn't make it into the methods section of a paper but determines whether your reaction works or fails.
Common Pitfalls That Cost Real Time and Money
Moisture is the most common problem, and everyone knows that. Lewis acids react with water. BF3·OEt2 fumes in air. AlCl3 hydrolyzes violently. But the subtler moisture issue is trace water in solvents that hasn't been fully removed by molecular sieves or distillation. You can run your reaction under argon with glassware dried in an oven and still have ten percent conversion because your solvent contained five hundred ppm of water. That water competes with your substrate for the Lewis acid and forms inactive hydroxo or oxo species. Karl Fischer titration of your solvents before use prevents this. It takes ten minutes and saves hours of failed reactions. Another frequent mistake is assuming that a stronger Lewis acid always gives a better result. Stronger doesn't mean better. A very strong Lewis acid may bind too tightly to your product, making isolation difficult or impossible without harsh workup conditions that degrade your compound. Mild Lewis acids often give cleaner reactions and easier purification. This is why catalytic amounts of less aggressive acids like Yb(OTf)3 or In(OTf)3 have gained popularity in recent years. They activate the substrate adequately and release the product more readily after the reaction completes. Product inhibition is a real issue that people overlook. When your Lewis acid coordinates to the product of your reaction as strongly as it coordinated to the starting material, the reaction slows down or stops entirely. This is particularly common when your product contains a free hydroxyl or amine group. The Lewis acid binds to that group and is no longer available to catalyze additional turnover. Adding a protective group or using a stoichiometric excess of Lewis acid can solve this, but both options have drawbacks. Protection adds steps. Excess Lewis acid complicates workup. The best approach is usually choosing a Lewis acid with lower affinity for your product functional groups in the first place.
Get the Full Details
Workup and Isolation Challenges
Removing Lewis acids from your product is often harder than running the reaction. Standard aqueous workup with dilute acid or base doesn't always work. AlCl3 forms gelatinous aluminum hydroxide precipitates that trap your product and make phase separation difficult. Boron compounds can form boric acid gels. Titanium residues are notoriously difficult to remove completely and show up as contaminants in NMR spectra even after multiple washes. The practical solution depends on your Lewis acid. For aluminum salts, a careful pour onto ice-cold dilute HCl helps dissolve the precipitate without emulsifying everything. For boron compounds, warm aqueous sodium bicarbonate usually converts boron species into water-soluble borates. For titanium, celite filtration through a short pad can remove a significant portion of the metal before you even attempt extraction. I also keep a supply of Chelex resin for polishing reactions where trace metal contamination is unacceptable for downstream steps. One workaround I rely on frequently is using immobilized Lewis acids on solid supports. Polymer-bound boron or titanium catalysts let you simply filter off the metal after the reaction completes. The tradeoff is usually lower activity compared to the homogeneous equivalents, and the cost of the supported reagent is higher per millimole. But when purification is the bottleneck, the immobilized option pays for itself quickly. I've had cases where a straightforward filtration replaced a six-hour workup and column chromatography.
Advanced Nuances Most People Miss
Lewis acid-catalyzed reactions often proceed through transition states that differ fundamentally from the uncatalyzed pathways. This means selectivity can change, not just rate. A reaction that gives predominantly the anti product without a Lewis acid might flip to syn selectivity with one present, because the Lewis acid changes the geometry of the transition state. This isn't speculation. It's documented extensively in aldol and Diels-Alder chemistry. The implication is that you can't simply substitute a Lewis acid into an existing protocol and expect the same selectivity profile. You need to evaluate stereochemical outcomes specifically for the catalyzed conditions. Counter-ion effects are another underrated factor. The anion associated with your Lewis acid matters more than most protocols acknowledge. Triflate anions are weakly coordinating and leave the Lewis acid center more exposed, which increases reactivity. Chloride and bromide are more strongly coordinating and can partially block the active site. This is why Al(OTf)3 is significantly more reactive than AlCl3 in many transformations despite having the same central metal. The counter-ion isn't a spectator. It's an active participant in determining catalytic activity and selectivity. There's also the question of whether your Lewis acid is acting as a true catalyst or a stoichiometric reagent. Many reactions reported as "catalytic" in the literature actually require near-stoichiometric amounts because of product inhibition, side reactions consuming the Lewis acid, or reversible binding that limits turnover. If your yield doesn't scale proportionally with catalyst loading at low loadings, you may be hitting a practical ceiling rather than a genuine catalytic cycle. Running a simple loading study from five mole percent to one hundred mole percent tells you immediately whether you're dealing with a true catalyst or a stoichiometric activator masquerading as one.
When Lewis Acids Are the Wrong Tool
Lewis acid catalysis has clear limitations. It doesn't help reactions that don't involve electrophilic activation. If your transformation requires nucleophilic catalysis or radical pathways, a Lewis acid won't help and may actively interfere. Brønsted acid catalysis is often a better choice for proton-transfer mechanisms. Organocatalysis avoids metal contamination entirely and is preferable when your product will be used in pharmaceutical applications where metal residue limits are strict. Thermally sensitive substrates are another case where Lewis acids can be problematic. Some Lewis acids require elevated temperatures to achieve meaningful activity, and those temperatures may exceed the stability window of your molecule. In those situations, switching to a milder Lewis acid or a different catalytic strategy entirely is more efficient than optimizing temperature and concentration. I've seen people push reactions to 120 degrees with AlCl3 when a simple phase-transfer catalyst at room temperature would have given better yields in less time. The cost and handling requirements of some Lewis acids also make them impractical for scale-up. AlCl3 requires careful quenching procedures that generate large volumes of aluminum salt waste. Boron trifluoride is a gas that requires specialized equipment for controlled addition. Some of the more exotic rare-earth triflates are expensive enough that the reagent cost becomes a significant factor at kilogram scale. For industrial applications, the cheaper and safer alternatives like iron salts or simple mineral acids are often sufficient and much easier to handle at production scale.
The field keeps evolving. New Lewis acids with improved selectivity, milder reactivity, and easier removal are being reported regularly. Ytterbium and scandium triflates have become standard options where they weren't twenty years ago. Session-specific ionic liquids as both solvent and Lewis acid source are an emerging area that could simplify workup considerably. But the fundamentals haven't changed. Understanding how your Lewis acid interacts with your substrate, your solvent, and your product is what determines whether the reaction works, not memorizing a list of recommended conditions from a paper.


