Understanding Resonance in Real Molecules
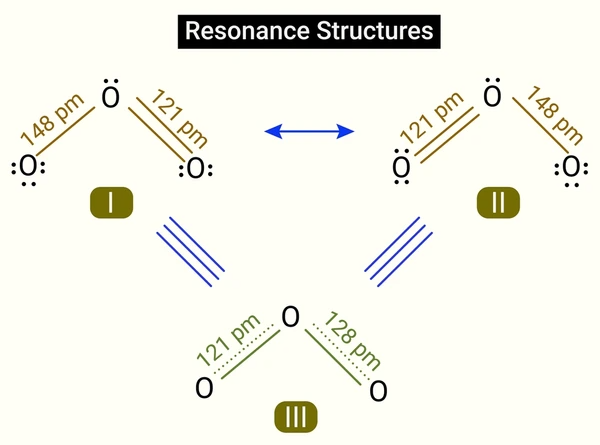
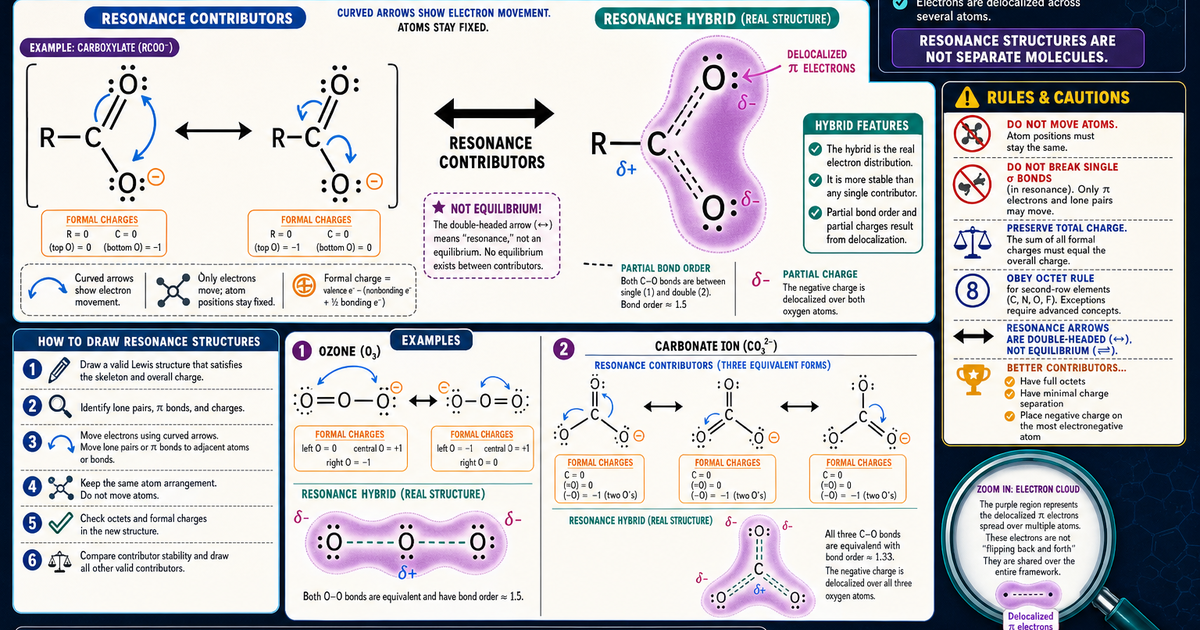
Resonance structures aren't real. They're a bookkeeping device chemists invented because the actual electron distribution in certain molecules can't be captured by a single Lewis structure. The molecule doesn't flip back and forth between drawings. It just exists as a hybrid, and the resonance structures are an imperfect way of describing that hybrid on paper. I learned this the hard way during my second year of organic chemistry, staring at ozone and convinced the oxygen atoms were literally switching places. They weren't. The molecule is symmetric. The two N-O bonds in nitrate are identical in length—124 picometers, right between a single and double bond. That's not because they're averaging over time. It's because electrons in those pi systems are delocalized across three atoms simultaneously.
Resonance Chemistry Demystified Unlocking The Hidden Electron Dance In Molecules
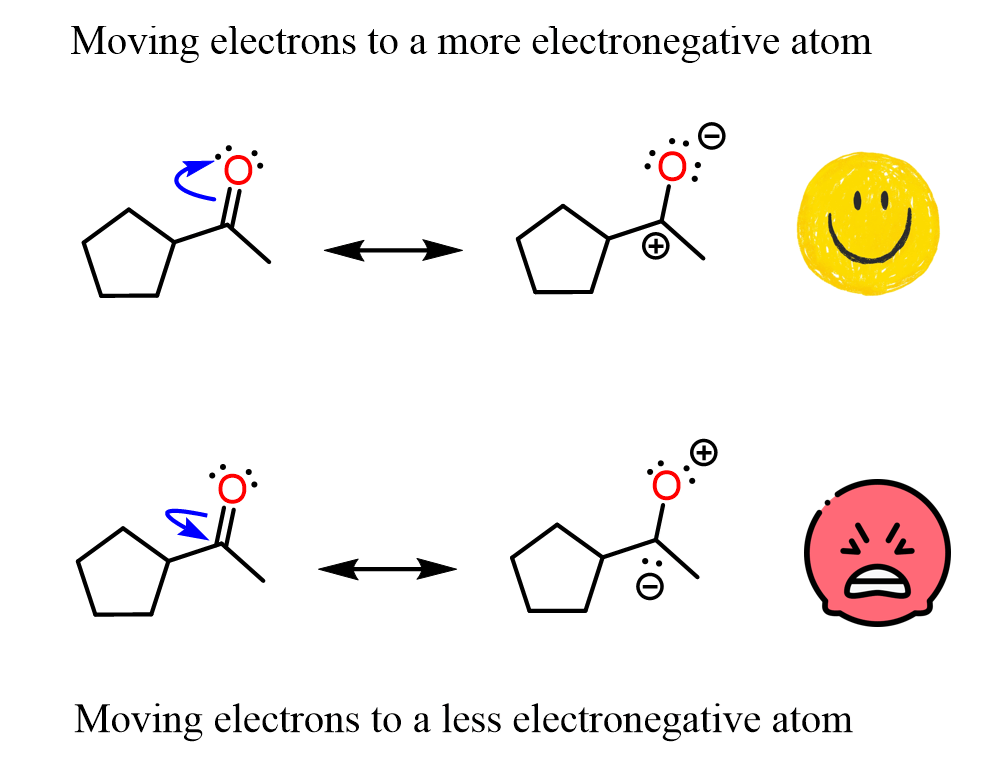
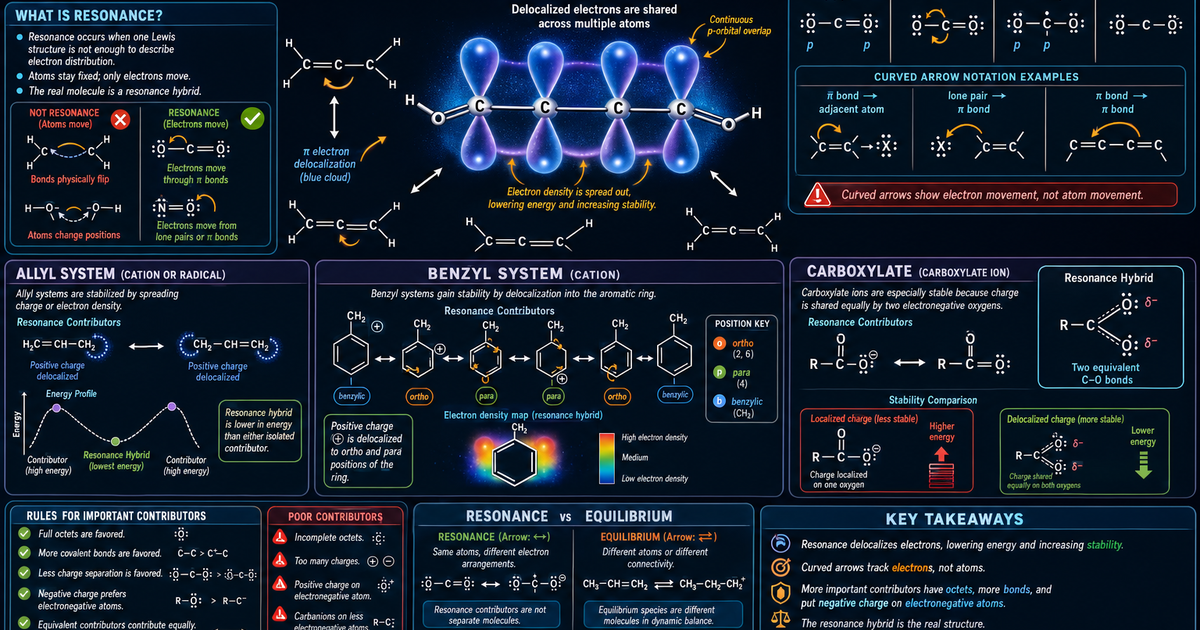
The practical process starts with identifying conjugated systems. Look for alternating single and multiple bonds, or a lone pair adjacent to a pi bond. When you see that pattern, you know delocalization is possible and you need to draw all reasonable contributing structures. The rules are straightforward but easy to mess up if you're rushing: Move only electrons. Atoms stay put. Any structure where bonds have broken or formed between different atoms isn't a resonance structure—it's a different molecule entirely, and it belongs in a reaction mechanism, not a resonance discussion. Electrons move in arrows from electron-rich areas to electron-poor areas. Pi electrons shift toward atoms or bonds that can better accommodate negative charge. Lone pairs donate into adjacent empty or partially filled orbitals. The big one people get wrong is handling charged species. Take the acetate anion. You draw the negative charge on one oxygen, then push a pi bond over and put the charge on the other oxygen. Both structures contribute equally. The real molecule has both oxygens bearing half a negative charge each. If you ever write that one oxygen is "more negative" in acetate, you've drawn it wrong. The carboxylate group is symmetric unless you've attached something that breaks the symmetry, like a substituent on one of the carbons.
I once spent three days debugging why my computational chemistry output didn't match literature values for the formate anion. The problem wasn't the software. It was me. I'd accidentally assigned different starting geometries to the two oxygen atoms during the initial input, which biased the wavefunction. The molecule has no reason to prefer one oxygen over the other, and the program was just faithfully reflecting my mistake back at me. I recentered everything and the results converged immediately.
Get the Full Details

When Resonance Actually Matters
Not every conjugated system benefits equally from resonance stabilization. The energy difference between drawing multiple resonance structures and just one is what we call resonance energy, and it varies wildly depending on the molecule. Benzene gains about 36 kilocalories per mole from delocalization. That's significant. It's why benzene doesn't undergo addition reactions the way cyclohexene does—it's too stable to give up that aromatic stabilization energy. But here's what most textbooks don't emphasize: resonance structures can sometimes be misleading about reactivity. Consider the enolate of acetone versus the enolate of acetylacetone. Both have resonance stabilization. But acetylacetone's enolate is stabilized by two carbonyl groups flanking the same carbon, and the resulting delocalization makes it roughly a thousand times more acidic than acetone. The pKa drops from about 19 to around 9. That's not a small difference. It's the difference between needing a strong base like sodium hydride to deprotonate and being able to use something as mild as hydroxide. The common pitfall is assuming all resonance contributors are equal. They almost never are. The major contributor is the one where every atom has a complete octet, where negative charge sits on the more electronegative atom, and where there are the fewest formal charges overall. Minor contributors still matter—they explain reactivity patterns that the major structure alone can't predict—but they contribute less to the actual hybrid.
Take the nitromethane anion, CH2NO2 minus. You might draw the negative charge on carbon and push electrons into the nitrogen-oxygen system. But the actual major contributor has the negative charge on oxygen, not carbon. Oxygen handles negative charge better. The carbon-centered structure isn't zero—there's still some carbanion character there, which is why the alpha hydrogens are somewhat acidic—but if you're predicting nucleophilic behavior based solely on a carbon-localized structure, your predictions will drift from what actually happens in solution.
Common Mistakes and How to Avoid Them
The most frequent error I see is violating the octet rule in second-row elements. You cannot draw a resonance structure where nitrogen has five bonds. Period. Some students will push electrons to create a pentavalent nitrogen and call it a day. It's not a valid resonance structure. It's just wrong. Nitrogen can expand its octet only when it's in row three or below. Sulfur can. Nitrogen cannot. Another error is treating resonance and tautomerism as the same thing. Tautomers involve actual proton transfer and represent different molecules in equilibrium. Keto-enol tautomerism is the classic example. The hydrogen physically moves from carbon to oxygen. Resonance structures have no atoms moving. Only electrons redistribute. If you need to break and form sigma bonds to interconvert your drawings, you're dealing with tautomers or conformers, not resonance. The hyphenation mistake matters too. Resonance hybrids are often depicted with dashed lines or partial bonds. A dashed line between two atoms means the bond order is fractional—1.5 in benzene, for instance. Don't draw that dashed line and also draw a full double bond in the same structure. That's contradictory notation. Pick one representation and stick with it consistently.

For molecules with heteroatoms, electronegativity differences become the dominant factor in determining which resonance form dominates. In amides, the nitrogen lone pair donates into the carbonyl pi system, giving the C-N bond partial double-bond character. That's why amide bonds are planar and why peptide bonds resist rotation. The resonance energy is about 17 to 20 kcal/mol. It's not as large as benzene's, but it's large enough to lock geometry in proteins and dictate the secondary structure of polypeptides. The barrier to rotation around a peptide bond is roughly 15 to 20 kcal/mol, which at room temperature means interconversion is slow on the NMR timescale but fast on the human timescale. That's why your protein unfolds when you heat it—the thermal energy overcomes the resonance stabilization that holds the backbone in place.
Practical Tools and Limitations
If you're doing this by hand, drawing all valid resonance structures for larger molecules gets tedious fast. A molecule like the bicarbonate ion has two major contributors. Something like a substituted phenoxide with multiple electron-donating and electron-withdrawing groups can have six or eight reasonable structures. Molecular orbital calculations through free software like ORCA or Gamess-US can map the actual electron density more accurately than any resonance drawing, but those require familiarity with computational chemistry input formats and take significantly longer to set up than a quick resonance analysis. For routine work, a spreadsheet-based resonance structure calculator or a web tool like ChemDraw's structure validation can catch invalid structures automatically—octet violations, impossible charges, atoms with incorrect valence. These tools won't tell you which structure is the major contributor, but they'll save you from wasting time on structures that violate basic quantum mechanics. The main limitation of the resonance model itself is that it breaks down for molecules where delocalization involves d-orbitals or where the electron distribution is better described by molecular orbital theory alone. Transition metal complexes are a good example. The resonance structure approach for organometallic compounds often produces nonsensical results because it can't properly account for back-bonding and the complex d-orbital interactions involved. For those systems, you're better off using crystal field theory or ligand field theory directly rather than forcing resonance drawings onto a problem they weren't designed to solve.
Another area where resonance predictions fail is in highly strained ring systems. Cyclopropenone has a dipole moment of about 5.1 Debye, which seems enormous until you consider the angle strain forcing the pi electrons into an unnatural distribution. The resonance contributor with a positive charge on the three-membered ring and a negative charge on the carbonyl oxygen is unusually important here, but only because the geometric constraints make the neutral structure energetically unfavorable. Standard resonance rules don't account for strain energy, so applying them without considering geometry gives misleading predictions about reactivity and stability. The practical takeaway is that resonance is a useful model when you know its boundaries. It works well for planar conjugated organic molecules with second-row elements. It gets shaky with non-planar systems, transition metals, and strained rings. In those cases, switch to molecular orbital theory or computational methods rather than trying to force the resonance framework into situations where it no longer provides accurate predictions.